Good Manufacturing Practice (GMP): History, Definition, and Importance
Good Manufacturing Practice (GMP) refers to a system of principles, procedures, and controls designed to ensure that products are consistently produced and controlled according to quality standards appropriate for their intended use. In the pharmaceutical sector, GMP governs every stage of manufacturing – from raw material sourcing and facility design to production, packaging, labeling, storage, and distribution. The primary goal of GMP is to ensure that products are safe, effective, and free from contamination. More than a set of operational guidelines, GMP represents a comprehensive quality assurance philosophy aimed at minimizing risks that cannot be eliminated solely through testing of the final product. These risks include contamination, mix-ups, deviations in potency, labeling errors, and inconsistencies in product quality. Today, GMP applies not only to pharmaceutical products and active pharmaceutical ingredients (APIs), but also to biologics, vaccines, diagnostics, food products, cosmetics, and medical devices.
Historical Development of GMP
The history of GMP is rooted in public health crises and the growing recognition that product safety cannot rely solely on end-product testing. Instead, quality must be built into the manufacturing process itself. The development of GMP has therefore been shaped by regulatory evolution, industrial growth, and lessons learned from catastrophic failures in pharmaceutical production over many years. The origins of GMP can be traced to the early twentieth century, when increasing industrialization of medicine highlighted the need for regulation. In the United States, the Pure Food and Drugs Act of 1906 marked one of the first major legislative efforts to prevent adulterated and misbranded drugs from entering the market. Although it did not establish GMP requirements, it laid the foundation for later quality regulations by recognizing the need for government oversight of drug manufacturing.
A major turning point occurred in 1937 with the Elixir Sulfanilamide tragedy, in which more than 100 people died after consuming a sulfanilamide preparation formulated with diethylene glycol, a toxic solvent. The disaster exposed severe gaps in manufacturing oversight and prompted the enactment of the Federal Food, Drug, and Cosmetic Act of 1938, which required manufacturers to demonstrate drug safety before marketing and granted regulatory authorities broader inspection powers. This event significantly influenced the philosophy that manufacturing processes, not just final products, must be regulated. Further momentum came from the thalidomide disaster in the late 1950s and early 1960s, when a sedative prescribed to pregnant women caused severe congenital deformities in thousands of infants worldwide. Although primarily associated with drug approval and clinical testing failures, the tragedy reinforced the necessity for stronger controls across the pharmaceutical lifecycle, including manufacturing. In response, regulatory reforms intensified globally, emphasizing quality systems and process controls.
Formal GMP regulations began to take shape in the 1960s. In 1963, the United States Food and Drug Administration (FDA) introduced the first GMP regulations, establishing legally enforceable requirements for manufacturing processes, sanitation, personnel practices, equipment, and recordkeeping. These regulations represented a paradigm shift from product testing alone to process-oriented quality assurance. The 1970s saw major global harmonization efforts. In 1969, the World Health Organization (WHO) published its first GMP guidelines, providing an international framework particularly valuable for countries developing pharmaceutical regulatory systems. WHO GMP became a foundational model for many national regulations and remains highly influential, especially in low- and middle-income countries.
As pharmaceutical science and manufacturing technologies advanced, GMP evolved accordingly. During the 1980s and 1990s, emphasis expanded beyond compliance with procedural rules toward robust quality systems. Concepts such as validation, qualification, documentation control, and change management became central components of GMP. Validation of manufacturing processes, analytical methods, cleaning procedures, and computerized systems became essential to demonstrate consistent product quality. A major milestone in modern GMP history was the emergence of international harmonization through the International Council for Harmonization (ICH) in the 1990s. ICH guidelines integrated scientific and risk-based approaches into GMP, promoting consistency across major regulatory jurisdictions. Around the same period, the principles of current Good Manufacturing Practice (cGMP) gained prominence, emphasizing that manufacturers must use up-to-date technologies and systems rather than relying on outdated compliance methods.
The twenty-first century has further transformed GMP through the incorporation of Quality by Design (QbD), risk management, data integrity principles, and continuous process verification. Modern GMP is no longer viewed merely as compliance with prescriptive rules but as a dynamic quality management system integrating science, risk assessment, and lifecycle control. Regulatory agencies such as the FDA, the European Medicines Agency (EMA), and WHO continue to revise GMP requirements to address emerging challenges, including biologics manufacturing, advanced therapies, sterile processing, digitalized systems, and globalized supply chains. The history of GMP reflects a progression from reactive regulation following public health disasters to a proactive and preventive framework focused on building quality into every stage of production (of medicines, foods, cosmetics, vaccines, medical devices and other goods).
Definition of GMP
Good Manufacturing Practice (GMP) can be defined as the part of quality assurance that ensures medicinal products are consistently produced and controlled to the quality standards appropriate to their intended use and as required by marketing authorization or product specifications. It encompasses all factors that influence product quality, including personnel competence, premises, equipment, materials, documentation, production operations, quality control testing, sanitation, and complaint handling. At its core, GMP is founded on the principle that quality cannot be tested into a product after manufacture; it must be designed and built into the process. This distinguishes GMP from quality control alone. While quality control focuses primarily on testing and verification, GMP covers the broader system that prevents errors before they occur.
Fundamental elements of GMP include proper documentation, validated processes, trained personnel, controlled environments, standardized operating procedures, traceability, and effective monitoring systems. Together, these elements ensure products meet identity, strength, purity, safety, and efficacy requirements consistently from batch to batch. Although GMP requirements may differ somewhat among regulatory jurisdictions, the central principles remain universal: prevent contamination, avoid mix-ups and errors, maintain consistency, and ensure that every product released meets established quality standards.
Importance of GMP
The importance of GMP lies primarily in protecting public health. Pharmaceutical products directly affect human life, and failures in quality can lead to therapeutic failure, toxicity, infection, or death. GMP provides the structured controls needed to prevent such outcomes by ensuring that products are manufactured under conditions that guarantee safety and reliability. One of the most important roles of GMP is contamination prevention. Particularly in sterile products and biologics, microbial, chemical, or particulate contamination can have catastrophic consequences. GMP establishes strict controls over environmental conditions, sanitation, personnel practices, and manufacturing operations to minimize these risks.
GMP is also essential for ensuring product consistency. Patients and healthcare providers rely on medicines to perform predictably. Variability in dosage strength, formulation, or stability can compromise efficacy and safety. Through validated processes and controlled operations, GMP ensures each batch is produced to the same predefined standards. Another critical importance of GMP is error prevention. Many product failures arise not from malicious intent but from preventable mistakes such as labeling errors, cross-contamination, or documentation failures. GMP reduces these risks through standardized procedures, training, traceability systems, and independent quality oversight.
From a regulatory perspective, GMP is fundamental for market authorization, inspection readiness, and international trade. Compliance demonstrates that a manufacturer operates according to recognized quality standards, facilitating product registration and access to global markets. For pharmaceutical companies, adherence to GMP is not only a legal obligation but also a strategic requirement for credibility and competitiveness. GMP is equally important for fostering consumer confidence. Patients trust that medicines are safe, effective, and manufactured under rigorous standards. This trust depends heavily on robust GMP systems. Failures in compliance can lead to recalls, regulatory sanctions, reputational damage, and loss of public confidence.
Economically, GMP supports efficiency by reducing deviations, rejects, rework, and recalls. While implementation requires investment, effective GMP systems reduce long-term costs associated with poor quality. In this sense, GMP is both a public health safeguard and a driver of sustainable pharmaceutical manufacturing. The importance of GMP extends beyond compliance. It embodies a preventive quality culture that prioritizes patient safety, product integrity, and continuous improvement. Its enduring relevance lies in the fundamental principle that medicines must not only be effective in theory but must be manufactured under conditions that ensure they are consistently safe and reliable in practice (Figure 1).
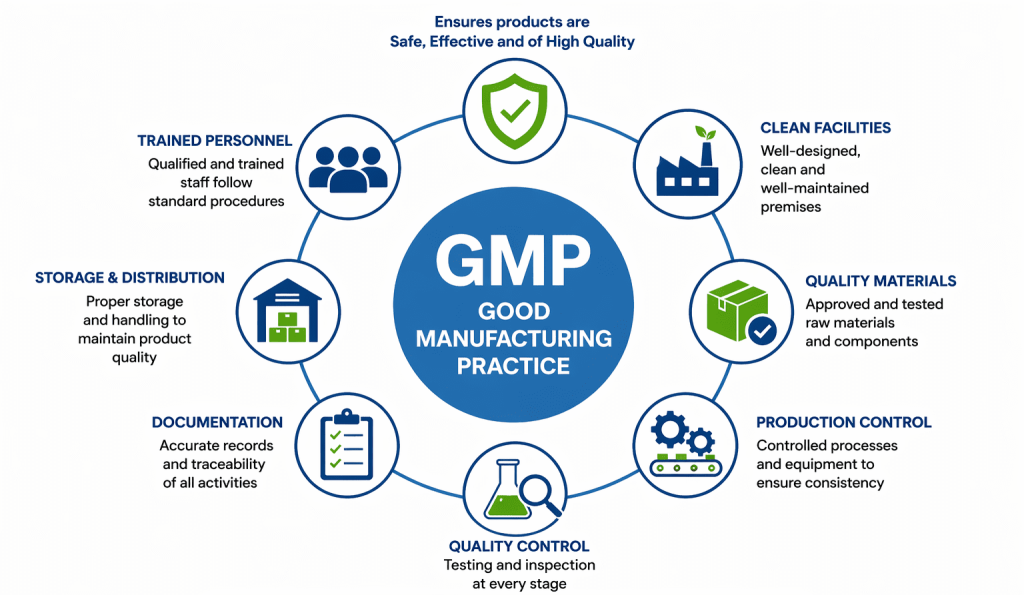
This figure illustrates the fundamental components of GMP and how they interact to ensure the production of safe, effective, and high-quality pharmaceutical products. It highlights key GMP pillars, including trained personnel, clean facilities, quality materials, production control, quality control, documentation, and proper storage and distribution, all of which collectively support product quality and patient safety. It shows that GMP is a comprehensive quality system designed to minimize risks such as contamination, errors, and inconsistencies throughout the manufacturing process.
Relationship Between GMP, quality assurance (QA), and quality control (QC)
Good Manufacturing Practice (GMP) provides the fundamental framework through which pharmaceutical products are consistently manufactured and controlled to meet predefined quality standards. Central to the successful implementation of GMP are two closely related but distinct components: Quality Assurance (QA) and Quality Control (QC). Although these concepts are often used interchangeably, they represent different but complementary pillars within pharmaceutical quality management. Together, QA and QC form the operational backbone of GMP, ensuring that products are not only manufactured correctly but also verified to meet safety, efficacy, and purity requirements before reaching patients.
Understanding the relationship between GMP, QA, and QC
GMP is a broad regulatory and operational system designed to minimize the risks inherent in pharmaceutical production that cannot be eliminated through testing of final products alone. These risks include contamination, mix-ups, deviations, and failures in consistency. Within this system, QA and QC function as integrated mechanisms that support the achievement of quality objectives.
Quality Assurance (QA) is the broader concept and can be defined as that part of GMP concerned with providing confidence that products will consistently meet the required quality standards appropriate for their intended use. QA encompasses all planned and systematic activities necessary to ensure quality is built into a product throughout its lifecycle. It is preventive in nature, focusing on designing systems and procedures that avoid defects before they occur. In essence, QA ensures that the right processes are in place to produce quality products.
Quality Control (QC), on the other hand, is a narrower component within QA and refers specifically to the part of GMP concerned with sampling, specifications, testing, documentation of results, and release procedures that verify whether products meet established quality criteria. QC is largely detective in nature, identifying defects or deviations through analytical and laboratory-based evaluations. While QA builds quality into the process, QC confirms that the output conforms to that quality.
This distinction is often summarized by the principle that QA is process-oriented, while QC is product-oriented. However, both are interdependent and inseparable under GMP. Without QA, there would be no structured systems to ensure quality is embedded in operations; without QC, there would be no scientific verification that those systems are producing acceptable products.
Quality assurance as a core element of GMP
QA encompasses all aspects that influence the overall quality of a finished pharmaceutical product. It extends beyond manufacturing operations and includes organizational structures, responsibilities, procedures, documentation systems, training, validation, audits, and product release processes. QA ensures that every activity affecting product quality is planned, controlled, monitored, and continuously improved. A critical function of QA under GMP is documentation control. Accurate and complete documentation forms the basis of traceability, accountability, and reproducibility in pharmaceutical manufacturing. Standard operating procedures (SOPs), batch manufacturing records, validation protocols, change controls, and deviation reports are all managed within the QA system to ensure consistency and compliance.
Another major role of QA is oversight of product release. Under GMP, products should not be released for distribution until authorized personnel have confirmed that they were manufactured and tested in accordance with approved procedures and specifications. This release process is a key QA responsibility and represents a critical safeguard for patient safety. Validation is another cornerstone of QA. Through process validation, cleaning validation, equipment qualification, and analytical method validation, QA ensures that systems and processes consistently perform as intended. This preventive approach reflects the fundamental GMP principle that quality cannot be tested into a product after manufacture but must be designed into every stage of production. QA also includes deviation management, corrective and preventive actions (CAPA), internal audits, risk management, and supplier qualification. These systems allow organizations to identify weaknesses, correct deficiencies, and prevent recurrence, thereby fostering continuous improvement and a culture of quality.
Quality control within the GMP framework
While QA establishes the systems that govern quality, Quality Control focuses on verifying compliance with established standards through scientific testing and analytical evaluation. QC is responsible for ensuring that raw materials, in-process samples, packaging materials, and finished products meet predefined specifications before use or release. Sampling and testing are central QC activities. Raw materials are analyzed to confirm identity, purity, and suitability before entering production. In-process testing monitors critical quality attributes during manufacturing, while finished product testing verifies characteristics such as potency, sterility, dissolution, stability, and microbial limits.
QC laboratories operate under strict GMP requirements related to equipment calibration, method validation, reference standards, analyst training, and documentation. These controls ensure reliability, accuracy, and integrity of test results. Through these activities, QC serves as a scientific checkpoint within the broader GMP system. QC is not limited to testing alone. It also includes specification development, out-of-specification investigations, environmental monitoring, and support for stability programs. These functions contribute to ongoing assurance that products remain safe and effective throughout their shelf life. Although QC often detects defects after they arise, its role within GMP extends beyond detection. QC data can provide critical feedback that supports process improvements, trend analysis, and risk reduction, reinforcing its integration with QA.
GMP as the foundation for product quality and patient safety
GMP guidelines outline the essential aspects of production and testing that influence product quality, ensuring that finished products are fit for consumption and free from microbial, chemical, and physical contamination. These guidelines cover all elements of pharmaceutical production, including personnel, facilities, sanitation, equipment, documentation, manufacturing operations, quality systems, and laboratory controls. The core objective of GMP as aforesaid is safeguarding patient health. Because pharmaceutical products directly affect human well-being, errors in manufacturing can have serious or even fatal consequences. GMP seeks to prevent such risks by requiring robust systems that assure consistency, purity, efficacy, and safety.
Recognizing this importance, many countries have made GMP compliance a legal requirement for pharmaceutical and medical device manufacturers. Regulatory agencies such as the US FDA, the EMA, and the WHO have established GMP guidelines that, while varying in detail, share common principles centered on quality assurance and risk prevention. These regulations reflect a global consensus that product quality cannot depend solely on end-product testing but must be controlled through well-designed and well-managed manufacturing systems. This is precisely where the relationship between GMP, QA, and QC becomes most significant.
Importance of integrating QA and QC under GMP
The effectiveness of GMP depends on the seamless integration of QA and QC. QA provides the preventive systems and governance structures needed to avoid defects, while QC supplies the analytical verification needed to confirm quality. Together, they create a comprehensive framework that supports compliance, consistency, and patient protection. This integration is essential not only for meeting regulatory requirements but also for maintaining public trust in pharmaceutical products. Failures in either QA or QC can lead to contamination events, recalls, regulatory sanctions, and risks to patient safety. Conversely, strong QA and QC systems support operational efficiency, reduce product failures, and promote continuous improvement. GMP, supported by robust QA and QC systems, represents far more than regulatory compliance. It is a quality philosophy centered on prevention, control, and accountability. Its principal purpose is to ensure that every pharmaceutical product released to the market is safe, effective, and manufactured to the highest standards, thereby fulfilling the fundamental objective of protecting patient health.
GMP in contamination control and documentation
GMP is a cornerstone of pharmaceutical quality systems, established to ensure that medicines, medical devices, and active pharmaceutical ingredients (APIs) are consistently produced and controlled according to appropriate quality standards. One of the primary objectives of GMP is to guarantee that pharmaceutical products are safe, effective, and of the required quality for patient use. Through a structured system of regulations, procedures, and controls, GMP ensures that quality is built into every stage of production rather than relying solely on final product testing. A fundamental principle of GMP is that quality cannot be inspected into a product after manufacture; it must be embedded throughout the production process. For this reason, GMP practices require pharmaceutical manufacturers to adopt preventive systems that minimize risks associated with contamination, errors, mix-ups, and product variability. In doing so, GMP serves not only as a regulatory requirement but also as a practical framework for protecting public health.
GMP compliance as a mandatory requirement in pharmaceutical manufacturing
Compliance with GMP is a mandatory requirement in pharmaceutical manufacturing worldwide. Regulatory agencies require manufacturers to adhere to GMP standards because products manufactured under substandard conditions may be considered adulterated, regardless of whether the finished product appears acceptable during testing. In pharmaceutical regulation, adulteration refers not only to contamination or product defects but also to failure to manufacture under controlled and compliant conditions. This concept underscores a critical principle of GMP: product quality is determined not only by the characteristics of the finished product but also by the integrity of the processes used to manufacture it. A medicine produced under poorly controlled conditions may pose significant risks even if it passes final analytical testing. Consequently, compliance with GMP is essential for both legal authorization and patient safety. GMP provides manufacturers with guidelines and operational controls for managing all factors that may affect product quality, including personnel, premises, equipment, raw materials, environmental conditions, and documentation. Through these controls, manufacturers can effectively identify, prevent, and manage contamination risks that may arise during production.
Sources of contamination in pharmaceutical manufacturing
One of the most critical areas addressed by GMP is contamination control. Contamination poses a serious threat to product quality and can compromise safety, efficacy, and stability. Because pharmaceutical products are often intended for vulnerable patient populations, even minimal contamination can have severe consequences. The following are major sources of contamination in a pharmaceutical manufacturing plant:
- Raw Materials: Raw materials represent one of the earliest and most significant sources of contamination risk. Active ingredients, excipients, solvents, and processing aids may introduce microbial, chemical, or particulate contaminants if not properly sourced, tested, and handled. GMP requires that raw materials be obtained from approved suppliers, subjected to incoming quality control testing, and stored under controlled conditions before use. Supplier qualification, material traceability, and testing against predefined specifications are essential measures used to minimize contamination risks originating from raw materials. Water, a critical raw material in pharmaceutical manufacturing, requires particular attention. Pharmaceutical-grade water systems must be designed, monitored, and maintained to prevent microbial growth, endotoxin contamination, and chemical impurities. Under GMP, water systems are treated as critical utilities and are subject to stringent validation and routine monitoring requirements.
- Environmental and airborne contamination: Air supply is another major source of contamination in manufacturing facilities, particularly in sterile production environments. Airborne particulates, dust, microorganisms, and aerosols can contaminate products during processing if environmental controls are inadequate. GMP addresses this risk through controlled manufacturing environments, including cleanrooms, heating ventilation and air conditioning (HVAC) systems, pressure differentials, air filtration systems such as high-efficiency particulate air (HEPA) filters, and environmental monitoring programs. These controls are designed to maintain clean production conditions and prevent both microbial and particulate contamination.Routine environmental monitoring, including air sampling, surface monitoring, and particulate assessments, forms an important part of GMP compliance and supports early detection of contamination risks.
- Equipment and manufacturing premises: Manufacturing equipment and premises are also recognized as critical contamination sources. Poorly maintained equipment, inadequate cleaning procedures, or unsuitable facility design can introduce residues, cross-contamination, and microbial hazards into products. GMP therefore requires equipment qualification, preventive maintenance, cleaning validation, and sanitation programs to ensure equipment performs consistently and does not compromise product quality. Equipment must be designed to permit effective cleaning, minimize product carryover, and prevent contamination during processing.Similarly, facility design under GMP must support logical process flow, segregation of operations, and prevention of cross-contamination. Layouts should minimize the risk of mix-ups and provide suitable environmental conditions for different manufacturing operations.
- Personnel: Personnel are often considered one of the most significant sources of contamination in pharmaceutical manufacturing. Human operators can introduce microorganisms, particulates, and errors into production processes through improper practices or inadequate training. For this reason, GMP places strong emphasis on personnel hygiene, gowning procedures, training, and competency. Employees involved in production must understand contamination risks and follow standardized procedures designed to maintain product integrity. Training is particularly important because even well-designed systems can fail if personnel do not implement them correctly. GMP therefore views trained and competent personnel as a critical component of contamination control.
- Packaging and storage-related contamination: Packaging materials can also contribute to contamination if they are improperly handled, unsuitable for product protection, or contaminated prior to use. GMP requires packaging materials to be tested, approved, and controlled to ensure they do not compromise product quality.Similarly, finished products may become contaminated or degraded during storage if environmental conditions such as temperature, humidity, and handling are not adequately controlled. GMP includes detailed requirements for warehousing, distribution, and storage conditions to preserve product quality throughout the supply chain.
Role of quality personnel in contamination prevention
Given the wide range of contamination risks, pharmaceutical microbiologists and quality assurance and quality control personnel play a critical role in GMP implementation. Their responsibility extends beyond detection of contamination to include development and oversight of preventive systems that control contamination at its source. These professionals are involved in environmental monitoring, microbial risk assessment, validation studies, deviation investigations, and implementation of corrective and preventive actions (CAPA). Their expertise supports the identification of critical contamination points and the development of sustainable control strategies. Contamination control under GMP is not a one-time activity but a continuous process requiring monitoring, risk assessment, and ongoing improvement. Quality personnel are central to maintaining this proactive approach.
Documentation as a fundamental principle of GMP
Another essential aspect of GMP is documentation, often summarized by the principle: “If it is not documented, it did not happen.” Documentation provides evidence that manufacturing activities were performed according to approved procedures and serves as the foundation for traceability, accountability, and regulatory compliance. Proper documentation and recordkeeping are indispensable to GMP because they allow manufacturers to demonstrate control over processes and provide data necessary for review, investigation, and product release. Documentation systems under GMP include standard operating procedures (SOPs), batch manufacturing records, cleaning records, calibration logs, environmental monitoring reports, deviation reports, validation protocols, and quality control test results. These records must contain accurate, complete, and detailed information regarding all aspects of production and quality assurance. Documentation errors, omissions, or falsification can compromise product integrity and constitute serious GMP violations.
Importance of recordkeeping and traceability
Accurate recordkeeping ensures traceability throughout the manufacturing process. Every material, process step, test result, and personnel action should be documented in a way that allows reconstruction of a product’s history. This traceability is essential for batch review, product release, complaint investigations, and recalls when necessary. Without comprehensive records, manufacturers cannot effectively demonstrate compliance or investigate quality issues. GMP also requires documentation systems to be controlled, reviewed, and protected against unauthorized changes. Increasingly, this includes data integrity principles governing both paper-based and electronic records to ensure reliability and authenticity. Well-maintained records also facilitate regulatory inspections. During inspections, documentation serves as objective evidence that processes are performed consistently and according to GMP requirements.
GMP as a system for quality and patient protection
GMP practices extend beyond preventing contamination and maintaining records; they establish a comprehensive quality system designed to ensure that pharmaceutical products are consistently manufactured to the highest standards. By controlling contamination risks from raw materials, water, air, personnel, equipment, packaging, and storage, GMP safeguards product purity and reliability. Through robust documentation systems, it ensures accountability, traceability, and regulatory compliance. These practices are not merely administrative requirements but essential safeguards for patient health. The ultimate purpose of GMP is to protect the patient by ensuring that every medicine or medical product reaching the market is safe, effective, and of assured quality. In this way, GMP serves both as a regulatory framework and as a practical philosophy of preventive quality management one that remains fundamental to modern pharmaceutical manufacturing.
Regulatory oversight and global enforcement GMP
GMP is a legally enforced system governed by national and international regulatory authorities. Because pharmaceuticals, foods, medical devices, and related products directly affect human health, governments around the world have established regulatory bodies responsible for ensuring that manufacturers comply strictly with GMP requirements. These agencies provide oversight, establish standards, conduct inspections, enforce compliance, and take regulatory action where necessary to protect public health. The existence of these regulatory organizations reflects a universal recognition that product safety cannot be left solely to manufacturers. Independent oversight is essential to ensure that products intended for human use are consistently produced under conditions that minimize risks of contamination, variability, adulteration, and therapeutic failure. Through regulatory enforcement of GMP, these agencies help ensure that medicines and food products reaching the public are safe, effective, and of acceptable quality.
Role of regulatory agencies in GMP compliance
Regulatory agencies are empowered by law to oversee manufacturers involved in producing pharmaceuticals, biologics, medical devices, and food products. Their primary responsibility is to ensure that manufacturers comply with established quality standards and operate within frameworks designed to safeguard public health. These agencies develop and enforce GMP regulations, issue licenses and approvals, inspect manufacturing facilities, evaluate quality systems, investigate product defects, monitor adverse events, and enforce corrective actions where violations occur. In doing so, they serve as critical guardians of product quality and consumer safety. One of the central principles underlying regulatory oversight is that manufacturers must demonstrate ongoing compliance with GMP rather than merely claiming adherence. Regulatory agencies verify this compliance through inspections, audits, review of manufacturing records, product testing, and post-market surveillance activities. Through these mechanisms, regulators help ensure that manufacturers maintain robust systems for contamination control, documentation, quality assurance, and risk management throughout the product lifecycle.
Major regulatory agencies responsible for GMP oversight
- United States Food and Drug Administration (FDA): The Food and Drug Administration (FDA) is one of the most recognized regulatory authorities globally and plays a major role in enforcing GMP compliance in the United States. The FDA regulates pharmaceuticals, biologics, medical devices, food products, and other health-related products through legally binding standards, including current Good Manufacturing Practice (cGMP) regulations. The FDA conducts routine and risk-based inspections of manufacturing facilities to assess compliance with GMP requirements. These inspections may be scheduled or unannounced and involve review of facilities, procedures, records, production operations, and quality systems. When deficiencies are identified, the FDA may issue warning letters, import alerts, product recalls, consent decrees, or even facility shutdowns in severe cases. Through this regulatory framework, the FDA plays a critical role in ensuring that products manufactured in or imported into the United States meet appropriate standards of safety and quality.
- National Agency for Food and Drug Administration and Control (NAFDAC): In Nigeria, the National Agency for Food and Drug Administration and Control (NAFDAC) serve as the principal regulatory authority overseeing food, drugs, cosmetics, medical devices, chemicals, and packaged water.NAFDAC is responsible for enforcing GMP compliance among manufacturers operating in Nigeria, conducting inspections, registering products, monitoring quality standards, and taking enforcement actions against noncompliant companies. Its role is particularly significant in ensuring the safety of locally manufactured and imported products, preventing circulation of substandard or counterfeit medicines, and protecting the Nigerian population from unsafe products.
- Medicines and Healthcare Products Regulatory Agency (MHRA): In the United Kingdom, the Medicines and Healthcare products Regulatory Agency (MHRA) regulates medicines, medical devices, and blood products. The MHRA enforces GMP through licensing, inspections, compliance monitoring, and post-market surveillance. Its GMP inspectorate evaluates manufacturers against established standards and ensures that pharmaceutical products meet rigorous requirements before reaching patients.
Other International Regulatory Agencies
Many countries maintain their own regulatory authorities responsible for GMP oversight. Examples include:
- Therapeutic Goods Administration (TGA) in Australia
- Ministry of Food and Drug Safety (formerly KFDA) in South Korea
- Medicines Control Council (MCC), now succeeded by SAHPRA, in South Africa
- Agência Nacional de Vigilância Sanitária (ANVISA) in Brazil
- European Medicines Agency (EMA), which coordinates regulation across the European Union
Although these agencies may differ in structure or regulatory processes, they share a common objective: ensuring that products intended for human use are manufactured according to accepted GMP principles. Increasingly, these agencies collaborate internationally through harmonization initiatives and mutual recognition agreements, promoting consistent global standards for pharmaceutical quality.
Inspections and regulatory enforcement
A major tool used by regulatory authorities to enforce GMP compliance is inspection. Inspections provide direct assessment of whether manufacturing facilities are operating in accordance with regulatory expectations. These agencies could carry out both routine and unannounced inspections.
Routine and unannounced inspections
Regulatory inspections may be routine, risk-based, pre-approval, or triggered by specific concerns such as complaints, recalls, or adverse events. Some inspections are announced, while others may occur without prior notice to evaluate day-to-day compliance. Inspection findings often determine whether a facility remains licensed to operate or whether corrective actions are required.
During inspections, regulators review key aspects of GMP implementation, including:
- Facility design and sanitation
- Personnel training and hygiene
- Documentation and recordkeeping
- Equipment maintenance and validation
- Production controls
- Laboratory operations
- Environmental monitoring
- Contamination control systems
- Quality assurance and quality control practices
Regulatory actions for non-compliance to GMP guidelines or principles
Manufacturers that fail to meet GMP requirements may face serious consequences. Depending on the severity of deficiencies, regulatory actions may include:
- Warning letters or deficiency notices
- Product recalls
- Suspension or revocation of licenses to produce or manufacture
- Import bans
- Seizure of products
- Partial or full plant shutdown
- Legal sanctions or prosecution
These enforcement actions emphasize that GMP compliance is not optional but a mandatory condition for continued operation.
Importance of staying current with evolving GMP standards
GMP is not static. Regulatory expectations evolve continually in response to technological advances, emerging risks, scientific developments, and lessons learned from quality failures. For this reason, manufacturers must continuously update their operations to align with revised guidelines and regulatory expectations. Remaining compliant requires ongoing training, system improvements, validation updates, and monitoring of regulatory developments. Failure to adapt to changing GMP requirements can place companies at risk of inspection findings, enforcement action, and loss of market access. Therefore, maintaining GMP compliance is not simply about meeting current regulations but about sustaining a culture of continuous quality improvement.
Consequences of poor GMP compliance
Failure to adhere strictly to GMP principles can have severe consequences for both manufacturers and consumers. Some of the major impact include:
Impact on manufacturers: Noncompliance can lead to major financial and operational losses. Product recalls, regulatory sanctions, litigation, and reputational damage can significantly affect profitability and long-term sustainability. Where severe violations occur, manufacturing operations may be suspended or permanently shut down. Such closures can result in loss of revenue, bankruptcy, reduced investor confidence, and unemployment for workers.Low product quality or contamination can also erode consumer trust, reducing demand and damaging brand reputation.
Impact on consumers and public health: The consequences for consumers can be even more serious. Products manufactured under poor GMP conditions may be contaminated, ineffective, unstable, or otherwise unsafe. Contaminated pharmaceuticals or food products may expose users to pathogenic microorganisms such as Salmonellosis-causing Salmonella, Pseudomonas, Escherichia coli, or Klebsiella, among others. Such contamination may lead to infection, treatment failure, toxicity, or even death. When contaminated products reach large populations, disease outbreaks may occur, resulting in significant morbidity, mortality, and economic burdens associated with medical treatment and outbreak control. These risks illustrate why strict adherence to GMP is fundamentally a public health imperative.
GMP as a critical defense against contamination
Pharmaceutical products can be contaminated before, during, or after manufacturing if proper controls are absent. Given the multiple potential sources of contamination including raw materials, water, personnel, equipment, air systems, and storage conditions it is impossible to rely on chance or end-product testing alone. GMP provides the preventive framework needed to control these risks through validated processes, environmental controls, sanitation, documentation, and quality oversight. Although eliminating all microbial presence from manufacturing environments may be unrealistic, GMP seeks to reduce contamination risks to controlled and acceptable levels through scientifically designed systems. This preventive philosophy is one of the greatest strengths of GMP and a key reason why regulatory oversight remains indispensable.
References
Arora D.R (2004). Quality assurance in microbiology. Indian J Med Microbiol, 22:81-86.
Ashutosh Kar (2008). Pharmaceutical Microbiology, 1st edition. New Age International Publishers: New Delhi, India.
Axelsen P.H (2002). Essentials of antimicrobial pharmacology. Humana Press, Totowa, New Jersey, USA. Al-Jasser A.M (2006). Extended – Spectrum Beta – Lactamases (ESBLs): A Global Problem. Kuwait Medical Journal, 38(3):171-185.
Block S.S (2001). Disinfection, sterilization and preservation. 5th edition. Lippincott Williams & Wilkins, Philadelphia and London.
Denyer S.P., Hodges N.A and Gorman S.P (2004). Pharmaceutical Microbiology. 7th ed. Blackwell Publishing Company, USA.
Nally J.D (Ed.) (2007). Good manufacturing practices for pharmaceuticals. Sixth edition. Informa Healthcare USA, Inc, New York.
Discover more from Microbiology Class
Subscribe to get the latest posts sent to your email.