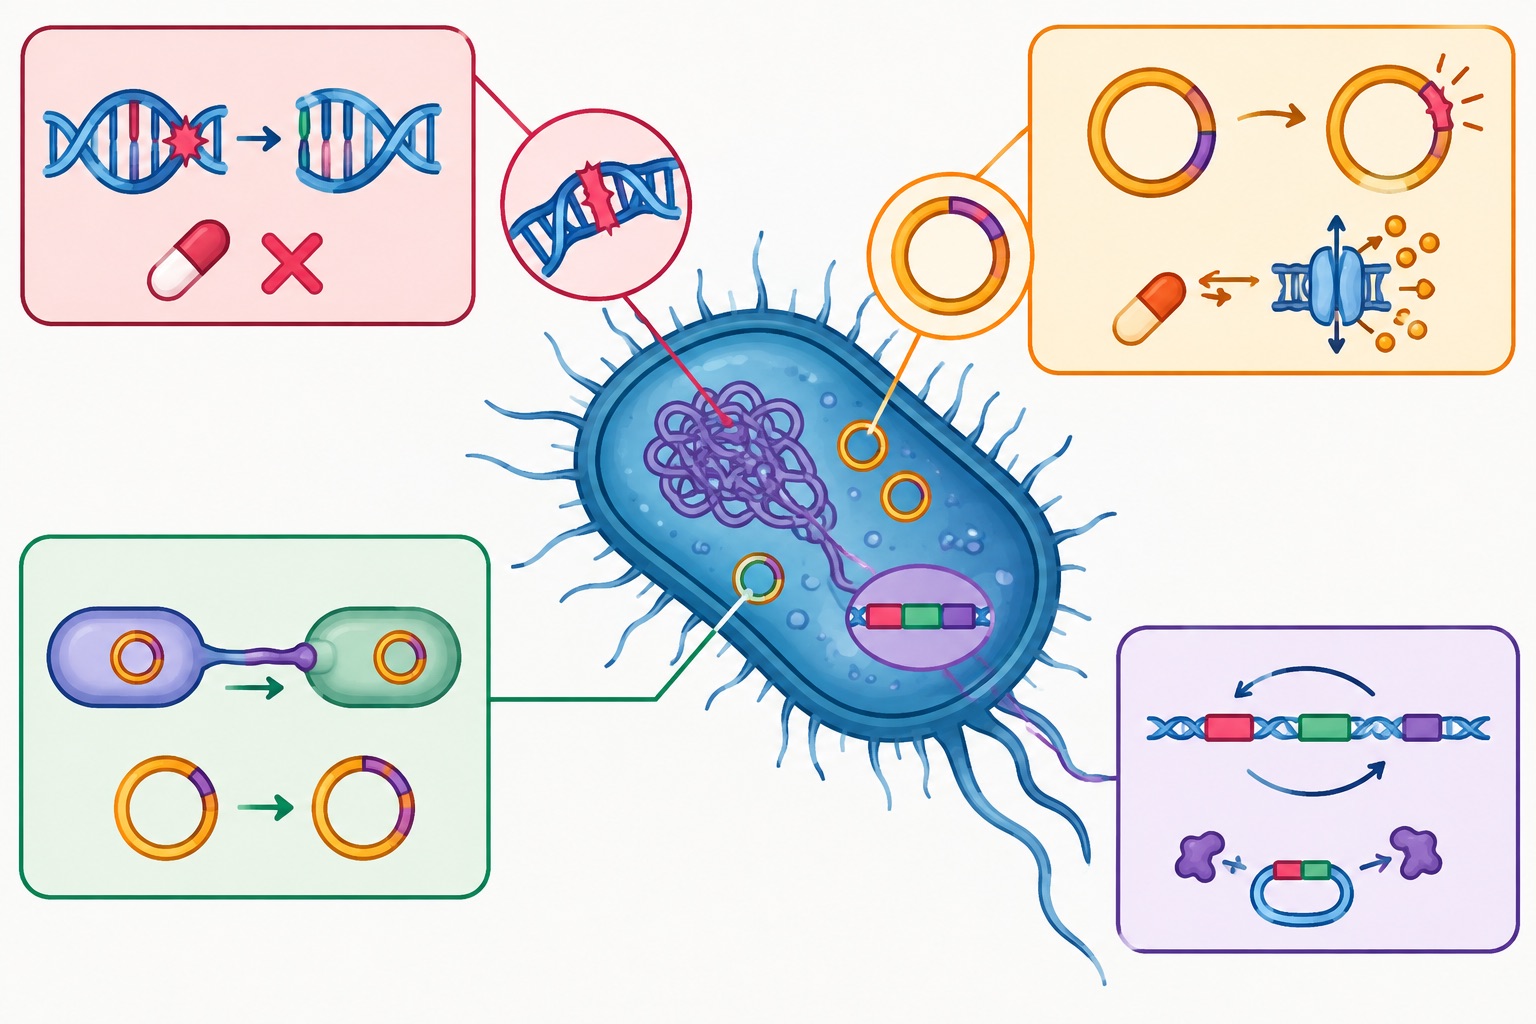
Antibiotic resistance is fundamentally a problem of microbial evolution in which genetic variation becomes translated into a measurable reduction in antimicrobial susceptibility. The phenomenon is not governed by a single resistance pathway but by a highly interconnected genetic architecture involving chromosomal mutations, horizontally acquired genes, mobile genetic elements, gene regulation, and adaptive changes in bacterial physiology. Microorganisms can therefore respond to antimicrobial exposure through several genetically distinct routes, ranging from alteration of an existing cellular protein to acquisition of an entirely new resistance determinant. This genetic flexibility enables bacterial populations to survive antibiotic concentrations that would otherwise inhibit growth or cause cell death.
The emergence of resistance should consequently be viewed as a population-level evolutionary process rather than simply as the appearance of a resistant phenotype in an individual bacterial cell. Within a microbial population, genetic variants arise continuously through replication errors, recombination, gene acquisition, and genome rearrangement. Antibiotic exposure then creates a selective environment in which susceptible cells are preferentially eliminated or inhibited, whereas variants carrying advantageous resistance traits survive and reproduce. Repeated exposure can progressively enrich these variants, particularly when antimicrobial use is inappropriate, unnecessary, poorly targeted, or insufficient to eliminate the infecting population. The resulting genetic enrichment may produce organisms with resistance to one antibiotic or, through accumulation and linkage of multiple determinants, multidrug-resistant phenotypes.
Antibiotic resistance has emerged as a complex genetic phenomenon in which microorganisms progressively acquire, modify, and regulate genetic traits that diminish the effectiveness of antimicrobial compounds. Although resistance is often recognized through observable treatment failure, its underlying biology is encoded within dynamic microbial genomes capable of rapid variation and adaptation. The genetic basis of resistance therefore extends beyond the presence of individual resistance genes; it encompasses mutation, gene regulation, genome plasticity, mobile genetic elements, and the exchange of genetic information between microorganisms.
At the genomic level, resistance can arise through spontaneous alterations in chromosomal DNA that modify antibiotic targets, metabolic pathways, membrane permeability, or regulatory networks. A single nucleotide substitution may alter the structure of a drug-binding site, while insertions, deletions, or larger genomic rearrangements can produce more substantial phenotypic changes. Importantly, resistance does not always depend on the acquisition of a completely new gene. Changes in promoter regions, transcriptional regulators, copy number, or signaling pathways can increase the expression of pre-existing cellular mechanisms and consequently alter antibiotic susceptibility. Such genetic changes demonstrate that resistance can emerge through functional reprogramming of the bacterial genome as well as through acquisition of foreign DNA.
The mobility of genetic information further expands the evolutionary potential of antibiotic resistance. Plasmids, transposons, insertion sequences, integrons, and bacteriophages can facilitate the movement of resistance determinants within and between microbial populations. Horizontal gene transfer enables genetic traits to cross taxonomic boundaries, allowing resistance-associated sequences to become incorporated into bacterial genomes that have not independently evolved the corresponding phenotype. This process can create multidrug-resistant lineages in which several resistance determinants are physically linked, increasing the potential for their simultaneous maintenance and dissemination under antimicrobial selection.
Another important genetic dimension is the interaction between resistance and bacterial fitness. Resistance-associated mutations may impose physiological costs by altering essential cellular processes, reducing growth efficiency, or disrupting regulatory balance. However, subsequent compensatory mutations can partially restore fitness while retaining the resistance phenotype. Resistant populations should not be viewed as genetically static entities. Their genomes may continue to evolve after resistance has emerged, generating variants with altered transmission potential, persistence, and antimicrobial response.
Antibiotic exposure provides a powerful selective environment in which genetic variants with increased survival can become enriched. Nevertheless, the evolutionary trajectory of resistance is influenced by factors beyond antibiotic concentration alone, including mutation rates, population structure, genetic linkage, ecological interactions, and the availability of transferable DNA. These interconnected processes create a genomic landscape in which resistance can emerge repeatedly, accumulate through multiple mechanisms, and persist even when selective conditions change.
Investigating the genetic basis of antibiotic resistance therefore requires an integrated perspective that connects DNA sequence variation with gene function, cellular physiology, and evolutionary dynamics. Advances in whole-genome sequencing, comparative genomics, transcriptomics, and molecular genetics now permit resistance-associated changes to be examined at increasingly precise levels. Such approaches can reveal not only which genetic alterations accompany resistance, but also how combinations of mutations and acquired determinants reshape microbial adaptation.
Chromosomal mutation and the evolution of intrinsic resistance
Chromosomal mutation represents one of the most direct genetic routes through which bacteria develop acquired resistance. Mutations may occur spontaneously during DNA replication or arise in response to cellular stress and environmental conditions. Although most mutations are neutral or deleterious, a small proportion can alter bacterial characteristics in ways that improve survival during antibiotic exposure. Such advantageous mutations may affect antibiotic-binding sites, membrane permeability, metabolic pathways, regulatory systems, DNA replication, or cellular structures required for drug activity.
A particularly important mechanism involves mutations within genes encoding antibiotic targets. Antibiotics exert their effects by interacting with specific bacterial molecules, such as ribosomal components, DNA-processing enzymes, cell-wall synthesis proteins, or metabolic enzymes. A nucleotide substitution that changes the amino-acid sequence or structural configuration of such a target can reduce antibiotic binding without completely eliminating the biological function of the protein. The antibiotic may therefore reach the bacterial cell but fail to interact efficiently with its intended target. Resistance in this context results not from removal of the drug but from a genetically modified cellular target with reduced drug affinity.
Mutations can also influence the quantity of an antibiotic target. Gene amplification or regulatory alterations may increase the production of cellular components affected by an antibiotic, effectively creating a greater functional reserve. In other circumstances, mutations alter metabolic pathways so that bacteria bypass the cellular process inhibited by the drug. These mechanisms illustrate that resistance can result from quantitative as well as qualitative genetic changes.
Regulatory mutations are particularly important because they can alter the expression of multiple genes simultaneously. Changes in transcription factors, promoters, repressors, or signal-responsive regulatory networks may increase the production of efflux proteins, modifying enzymes, membrane proteins, or stress-response components. A single regulatory mutation can therefore produce a broader resistance phenotype than a mutation affecting one structural protein. This illustrates why the genetic basis of resistance cannot be adequately understood by examining resistance genes alone; the regulatory context in which those genes operate is equally important.
Chromosomal resistance can also carry a biological cost. A mutation that changes a protein involved in antibiotic susceptibility may simultaneously reduce enzymatic efficiency, alter membrane function, or interfere with normal cellular metabolism. Bacterial populations can subsequently acquire compensatory mutations that reduce these fitness penalties while retaining resistance. Resistance evolution may therefore proceed through sequential genetic stages: initial acquisition of a resistance-conferring alteration followed by secondary mutations that improve physiological performance.
Horizontal gene transfer and the mobility of resistance determinants
Mutation is only one component of resistance evolution. Bacteria possess remarkable mechanisms for acquiring genetic information from other microorganisms, allowing resistance traits to spread without requiring independent mutation in every recipient population. Horizontal gene transfer (HGT) provides a genetic shortcut through which previously susceptible bacteria can acquire pre-existing resistance determinants.
Three major HGT mechanisms (conjugation, transformation, and transduction) contribute to horizontal movement of bacterial DNA. Conjugation commonly involves direct cell-to-cell transfer of plasmid-associated DNA and is particularly important because plasmids may carry several resistance genes simultaneously. Transformation involves uptake and incorporation of extracellular DNA. Transduction uses bacteriophages as vehicles for the movement of bacterial genetic material. The biological consequences of these processes extend beyond individual cells because successful transfer can introduce resistance determinants into a new bacterial lineage and subsequently permit their vertical inheritance.
Plasmids are especially important in the genetic epidemiology of antibiotic resistance because they can replicate independently of the bacterial chromosome. A plasmid carrying a resistance gene can therefore be maintained within a population and, under suitable conditions, transferred between bacterial cells. When several resistance genes occur on the same mobile element, selection by one antibiotic may indirectly preserve resistance to other antibiotics. This phenomenon, known as co-selection, can contribute to the persistence of multidrug-resistant populations even when exposure to every corresponding antibiotic is not continuous.
Transposons and insertion sequences further increase genomic flexibility. These mobile DNA elements can relocate genetic sequences between plasmids and chromosomes or facilitate the assembly of resistance-associated regions. Integrons represent another important genetic platform because they can capture and express gene cassettes, including antimicrobial resistance determinants. The combination of plasmids, transposons, insertion sequences, and integrons creates a dynamic genetic network in which resistance genes can be acquired, rearranged, linked, and disseminated.
The clinical significance of HGT lies in its capacity to accelerate resistance evolution. A bacterial population does not necessarily need to wait for a beneficial mutation to arise spontaneously if an appropriate resistance determinant is already present within its microbial environment. This capacity transforms the microbial community into a reservoir of genetic information from which resistance traits can potentially be recruited.
Molecular mechanisms produced by resistance-associated genetic changes
Genetic alterations ultimately produce resistance through changes in bacterial physiology. One major mechanism is enzymatic destruction or modification of the antibiotic. Resistance genes can encode enzymes capable of hydrolyzing, chemically modifying, or otherwise neutralizing antimicrobial compounds before they reach their cellular targets. β-lactamases provide a prominent example because they hydrolyze the β-lactam ring present in numerous antibacterial agents. Different β-lactamase families possess different substrate profiles, creating substantial variation in the range of antibiotics that can be compromised.
Another mechanism involves modification of the antibiotic target. Genetic changes may alter the target structure sufficiently to reduce drug binding while preserving essential cellular activity. Alternatively, bacteria may acquire genes encoding proteins that chemically modify the target itself. Target protection represents another strategy in which resistance-associated proteins interact with the antibiotic-target complex and reduce the inhibitory effect of the drug.
Reduced intracellular drug accumulation provides a different genetic solution. Bacterial membranes regulate the movement of molecules into and out of the cell, and genetic changes affecting membrane proteins can decrease antibiotic entry. Resistance genes may also encode active efflux systems that transport antibiotics from the cytoplasm or periplasm back into the extracellular environment. When influx is reduced and efflux is increased, the intracellular concentration of an antibiotic may remain below the level required for effective inhibition.
The genetic control of permeability and efflux is frequently interconnected. Mutations affecting porins can reduce antibiotic uptake, while regulatory mutations can increase efflux-pump expression. When these mechanisms occur simultaneously with enzymatic degradation or target modification, their effects may be additive or synergistic. Resistance phenotypes observed in clinical isolates may represent the combined output of several genetic mechanisms rather than the action of a single resistance gene.
Metabolic bypass provides another route to resistance. If an antibiotic blocks an essential biochemical pathway, bacteria may acquire mutations or genes that permit alternative metabolic routes. Similarly, amplification of metabolic enzymes or replacement of drug-sensitive proteins with functionally related but drug-insensitive variants can preserve cellular activity in the presence of an antibiotic. These mechanisms emphasize that resistance is fundamentally a problem of maintaining cellular function despite pharmacological interference.
Regulation, selection, and genetic adaptation under antibiotic exposure
The presence of a resistance determinant does not necessarily mean that it is continuously expressed at the same level. Bacteria possess sophisticated regulatory systems that determine when particular genes are transcribed and how strongly their products are produced. Resistance may therefore depend not only on the genetic sequence itself but also on its transcriptional and post-transcriptional regulation.
Antibiotic exposure can create selective conditions that favor cells carrying resistance-associated variants. When susceptible cells are inhibited, resistant cells experience reduced competition for nutrients and ecological space. Their descendants consequently represent an increasing proportion of the population. Repeated antimicrobial exposure can reinforce this process, particularly when treatment creates prolonged periods in which drug concentrations impose selection without completely eliminating the bacterial population.
This evolutionary process helps explain why inappropriate antibiotic use can contribute to resistance. Unnecessary prescriptions, inappropriate drug selection, incorrect dosing, prolonged exposure, and antimicrobial use outside clinical medicine can increase the ecological opportunities for resistant variants to persist. Antibiotic use in livestock and agriculture can also contribute to selection within microbial communities, creating reservoirs from which resistance determinants may potentially disseminate into other bacterial populations.
Antibiotic exposure does not simply select for pre-existing resistance. It can also alter the evolutionary environment in which new variants emerge and are maintained. Stress-response pathways, DNA repair processes, and changes in mutation dynamics may influence the generation of genetic diversity. The resulting population can contain multiple variants with different combinations of resistance determinants, creating a heterogeneous evolutionary landscape.
Resistance can also persist after the original selective pressure declines. This may occur when resistance genes impose little fitness cost, when compensatory mutations reduce their physiological burden, or when resistance determinants are genetically linked to other traits that remain advantageous. Withdrawal of a particular antibiotic does not necessarily result in immediate disappearance of the corresponding resistance phenotype.
Genomic plasticity, multidrug resistance, and emerging genetic landscapes
An increasingly important perspective is that antibiotic resistance should be studied as a property of a changing microbial genome rather than as an isolated gene–drug relationship. Bacterial genomes are highly plastic and can undergo mutation, recombination, gene acquisition, amplification, deletion, and rearrangement. These processes allow microorganisms to assemble complex resistance profiles in response to changing ecological and therapeutic pressures.
Multidrug resistance may develop when multiple resistance determinants accumulate within the same bacterial lineage or become physically linked on mobile genetic elements. In such circumstances, selection for one resistance trait can indirectly maintain several others. This genetic linkage complicates antimicrobial treatment because the phenotype reflects the combined activity of several independent or interacting mechanisms.
Modern genomic approaches provide new opportunities to investigate this complexity. Whole-genome sequencing can identify chromosomal mutations, plasmid-associated determinants, mobile elements, and previously unrecognized genetic variants. Comparative genomics can reveal differences between susceptible and resistant isolates, while transcriptomic approaches can determine whether resistance-associated genes are actively expressed under particular conditions. Functional genomic techniques can further distinguish genetic changes that merely correlate with resistance from those that directly contribute to the phenotype.
The better understanding of antibiotic resistance will therefore depend on integrating genotype, gene expression, cellular physiology, and evolutionary history. Such integration is particularly important for identifying emerging resistance mechanisms before they become widespread. Rather than viewing resistance as a fixed characteristic acquired at a single point in time, it is more informative to regard it as a continuously evolving genetic phenotype shaped by mutation, gene exchange, regulatory adaptation, ecological selection, and compensatory evolution. This perspective provides a stronger conceptual basis for surveillance, molecular diagnosis, antimicrobial stewardship, and the development of therapeutic strategies capable of addressing increasingly complex resistance architectures.
Cross-resistance and its genetic relationship to antibiotic resistance
Cross-resistance is a phenomenon in which a single genetic or physiological adaptation enables a microorganism to withstand several antimicrobial agents, even when those drugs differ in their chemical structures or clinical applications. Unlike resistance that develops independently against each antibiotic, cross-resistance can arise because multiple drugs share a cellular target, depend on similar pathways for entry, or are affected by the same resistance mechanism. It therefore represents an important genetic dimension of multidrug resistance and can significantly complicate antimicrobial treatment.
The genetic basis of cross-resistance commonly involves mutations or acquired genes that simultaneously influence the activity of several antibiotics. One important mechanism is target modification. A mutation in a chromosomal gene encoding a common antibiotic target may reduce the binding of multiple drugs that act on the same cellular structure. Similarly, acquisition of a resistance gene may produce an altered target protein that has reduced affinity for an entire class of antibiotics.
Efflux-pump systems provide another major genetic mechanism. Bacteria possess genes encoding membrane-associated transport proteins capable of removing antimicrobial compounds from the cell. Mutations in regulatory genes can cause constitutive or increased expression of these pumps. Because an individual efflux system may recognize structurally unrelated compounds, its overexpression can produce resistance to several antibiotics simultaneously. Thus, a single regulatory alteration can generate a broad cross-resistant phenotype rather than resistance to one specific drug.
Changes in cell-envelope permeability can also promote cross-resistance. Mutations that reduce the production or function of membrane porins may restrict the entry of multiple antibiotics into the bacterial cell. When reduced permeability occurs together with increased efflux, the intracellular concentration of several antimicrobial agents can fall below their effective levels.
Cross-resistance can additionally emerge through shared enzymatic mechanisms. Some resistance enzymes have broad substrate ranges and can modify or degrade several related antibiotics. β-lactamase genes, for example, may encode enzymes capable of compromising multiple β-lactam agents. The genetic acquisition of such determinants can therefore generate resistance across a group of therapeutically related compounds.
An important distinction is that cross-resistance is not synonymous with co-resistance. Cross-resistance occurs when one mechanism affects multiple antibiotics, whereas co-resistance refers to the presence of separate resistance determinants that are genetically linked, often on plasmids, transposons, or integrons. Both mechanisms can contribute to multidrug resistance, but their genetic origins are different.
From an evolutionary perspective, cross-resistance demonstrates how one genetic alteration can have consequences extending beyond a single antimicrobial agent. Selection imposed by one antibiotic may enrich bacterial variants carrying a mechanism that simultaneously reduces susceptibility to other antibiotics. Antimicrobial exposure can shape the frequency and distribution of resistance-associated genes even when the other affected drugs have not been directly used.
References
National Research Council (US) Committee to Study the Human Health Effects of Subtherapeutic Antibiotic Use in Animal Feeds. (1980). The effects on human health of subtherapeutic use of antimicrobials in animal feeds: Appendix C, Genetics of antimicrobial resistance. National Academies Press.
Nadeem, M. S., Rasool, R., Afzal, M., Ullah, I., Murtaza, B. N., Daoub, R. M. A., Chaieb, K., Alzarea, S. I., Kazmi, I., & Ashraf, G. M. (2026). Genes and physiological strategies in bacterial antibiotic resistance. Frontiers in Microbiology, 17, 1811959.
Harris, M., Fasolino, T., Ivankovic, D., Davis, N. J., & Brownlee, N. (2023). Genetic factors that contribute to antibiotic resistance through intrinsic and acquired bacterial genes in urinary tract infections. Microorganisms, 11(6), 1407.
Coleman, J. P., & Smith, C. J. (2007). Microbial resistance. In S. J. Enna & D. B. Bylund (Eds.), xPharm: The comprehensive pharmacology reference (pp. 1–3). Elsevier.
Franklin, T. J., & Snow, G. A. (1998). The genetic basis of resistance to antimicrobial drugs. In Biochemistry and molecular biology of antimicrobial drug action. Springer.
Hussain, H. I., Aqib, A. I., Seleem, M. N., Shabbir, M. A., Hao, H., Iqbal, Z., Kulyar, M. F.-e.-A., Zaheer, T., & Li, K. (2021). Genetic basis of molecular mechanisms in β-lactam resistant Gram-negative bacteria. Microbial Pathogenesis, 158, 105040.
Pontes, M. H., & Groisman, E. A. (2020). A physiological basis for nonheritable antibiotic resistance. mBio, 11, e00817-20.
Jian, Z., Zeng, L., Xu, T., Sun, S., Yan, S., Yang, L., et al. (2021). Antibiotic resistance genes in bacteria: Occurrence, spread, and control. Journal of Basic Microbiology, 61, 1049–1070.
Courvalin, P. (1994). Transfer of antibiotic resistance genes between Gram-positive and Gram-negative bacteria. Antimicrobial Agents and Chemotherapy, 38, 1447–1451.
Discover more from Microbiology Class
Subscribe to get the latest posts sent to your email.