Good laboratory practice (GLP)
Good laboratory practice (GLP) is a quality system designed to ensure the integrity, reliability, traceability, and verifiability of data generated in non-clinical health and environmental safety studies. It establishes a systematic framework for organizing laboratory activities, documenting experimental procedures, defining responsibilities, and maintaining accountability throughout the study lifecycle.
The primary objective of GLP is to ensure that regulatory decisions are supported by reliable, reproducible, and adequately documented scientific evidence. GLP is particularly important in the safety assessment of pharmaceuticals, pesticides, veterinary medicines, industrial chemicals, cosmetic ingredients, and environmental contaminants. It requires studies to be conducted in accordance with approved study plans, established standard operating procedures (SOPs), and applicable regulatory requirements.
The Organisation for Economic Co-operation and Development (OECD) has established internationally recognized GLP principles to promote the quality and mutual acceptance of non-clinical safety data among participating countries. In the United States, the Food and Drug Administration (FDA) enforces GLP regulations for specified non-clinical laboratory studies, while European regulatory frameworks incorporate applicable GLP requirements into the evaluation of safety data. Although regulatory requirements may vary by jurisdiction and study type, their common elements include qualified personnel, standardized procedures, independent quality assurance, appropriate equipment maintenance and calibration, controlled documentation, and secure record retention.
A fundamental principle of GLP is its emphasis on data integrity rather than predetermined experimental outcomes. GLP does not establish whether a substance is safe or hazardous; rather, it ensures that the evidence used to reach such conclusions is generated, recorded, analyzed, and reported through controlled, transparent, and auditable processes. All relevant findings, including unexpected observations, adverse effects, deviations from approved procedures, and negative results, must be appropriately documented and reported.
GLP promotes scientific accountability by clearly defining personnel responsibilities, controlling and documenting procedural changes, and maintaining comprehensive records from study initiation through final reporting and archiving. Beyond regulatory compliance, GLP provides a robust foundation for evaluating non-clinical safety data, facilitating comparisons across studies and laboratories, and supporting evidence-based regulatory decisions. GLP implementation helps safeguard the quality of scientific evidence used to protect human health and the environment.
GLP organizational structure, study management, and personnel responsibilities
The organizational structure of GLP establishes a clear allocation of authority and responsibility for the conduct of regulated non-clinical studies. Unlike quality systems organized primarily around manufacturing batches, GLP frameworks are generally study-centered, with each study conducted in accordance with a predefined protocol and supported by documented SOPs. This structure enables regulatory authorities to reconstruct the course of an investigation, identify the personnel responsible for specific activities, verify the methods employed, and assess the traceability of reported conclusions to the original study data.
The Study Director occupies a central role within this framework and bears overall responsibility for the conduct of the study in accordance with GLP principles. Key responsibilities include ensuring adherence to the approved protocol and applicable SOPs, assessing and documenting deviations, coordinating the contributions of study personnel, and overseeing the collection, analysis, and interpretation of study data. The Study Director must also ensure that the final report accurately reflects the study’s methods, results, and conclusions. For studies involving multiple sites, effective communication, clearly defined responsibilities, and appropriate documentation are essential to maintaining consistency, continuity, and accountability across locations.
Study personnel are responsible for performing assigned tasks competently, following applicable procedures, and recording observations and results promptly, accurately, and contemporaneously. Their activities may include specimen preparation, animal care and observations, analytical measurements, sample processing, equipment operation, and electronic data entry. Personnel must possess appropriate qualifications, experience, and training for their assigned duties, including task-specific competency where required. Documented training records, qualifications, and evidence of relevant experience provide assurance that study activities are performed by suitably competent individuals and support the reliability and traceability of the resulting data.
Test facility management is responsible for establishing and maintaining the organizational and operational conditions necessary for GLP compliance. This includes providing adequate personnel, suitable facilities, properly maintained equipment, appropriate materials, documented procedures, and effective systems for data and record management. Management must ensure that responsibilities and reporting relationships are clearly defined, that personnel receive appropriate training, and that the organizational structure supports the independence and effectiveness of quality assurance activities. These responsibilities establish the infrastructure within which individual studies can be conducted consistently and in accordance with applicable GLP requirements.
The Quality Assurance Unit (QAU) performs an independent monitoring function to assess whether studies and associated activities comply with applicable GLP principles, approved protocols, and relevant procedures. Its responsibilities may include conducting facility-based, process-based, and study-based inspections; examining study records and procedures; and verifying that final reports accurately reflect the reported methods and results. The QAU communicates inspection findings to the Study Director, relevant personnel, and management, as appropriate, so that identified deficiencies can be evaluated and addressed through documented corrective actions. The scope and frequency of inspections should be appropriate to the nature and progress of the activities being monitored.
The independence of quality assurance is a fundamental element of the GLP framework rather than a procedural formality. By separating compliance monitoring from the direct conduct of experimental activities, the QAU provides an objective assessment of whether established requirements are being followed. This independent oversight does not transfer or diminish the Study Director’s responsibility for the overall conduct of the study, nor does it relieve management of its responsibility to provide the resources and organizational support necessary for compliance.
Together, management, the Study Director, study personnel, and the QAU establish a system of complementary and clearly defined responsibilities that supports documented decision-making, data traceability, and objective evaluation of study conduct. This organizational framework enables regulatory authorities to assess not only the reliability of the reported findings but also the adequacy of the processes through which those findings were generated.
GLP experimental protocols, laboratory operations, and data integrity
The operational foundation of GLP is the controlled conduct of studies in accordance with documented and approved experimental protocols. A protocol defines the study objectives, design, test and reference materials, experimental methods, relevant measurements, and planned analytical procedures. Depending on the nature of the investigation, it may also specify test-system selection, dose levels, observation schedules, sampling procedures, statistical methods, and criteria for evaluating results. Establishing these elements before experimental work begins minimizes discretionary decision-making, promotes consistency, and provides a defined basis for assessing whether the study was conducted as intended.
SOPs complement the study protocol by establishing standardized, reproducible methods for routine laboratory activities. These procedures may address equipment operation, sample identification, reagent preparation, specimen handling, cleaning, instrument maintenance, data recording, and archiving. SOPs must be appropriately authorized, readily accessible to relevant personnel, and maintained through controlled revision processes. Changes to established procedures should be documented and communicated to ensure that personnel use the applicable version and understand any implications for laboratory operations or study conduct.
GLP also requires appropriate control of laboratory facilities, equipment, reagents, test systems, and study materials. Instruments used to generate critical measurements must be maintained, calibrated, and operated in accordance with documented procedures and predefined acceptance criteria, as applicable. Calibration and maintenance records provide evidence of equipment performance and help establish whether instruments were suitable for their intended use when measurements were obtained. Test and reference materials must be appropriately identified, handled, stored, and characterized, where required. Clear sample labelling and documented chain of custody support traceability and help prevent misidentification, loss, or mix-ups during transfers between personnel, laboratory areas, or analytical facilities.
Data integrity is a fundamental principle of GLP, ensuring that study records remain reliable, traceable, and suitable for independent review. Original observations and measurements must be recorded promptly and accurately, with sufficient context to preserve their meaning and support subsequent verification. Relevant records may include laboratory notebooks, instrument-generated files, chromatograms, photographs, electronic records, calculations, environmental monitoring results, and associated documentation. Corrections must be attributable, legible, and appropriately justified, while preserving the original entry and the history of any subsequent changes. Original records must not be silently overwritten, obscured, or deleted.
The ALCOA principles (Attributable, Legible, Contemporaneous, Original, and Accurate) provide a widely used framework for evaluating data integrity. Under this framework, records must identify who performed an activity or recorded data, remain readable, be documented at the time of the activity, preserve the original record or a verified true copy, and accurately reflect the observation or result. Extended interpretations, commonly described as ALCOA+, additionally emphasize completeness, consistency, enduring preservation, and availability throughout the required retention period. For electronic records, appropriate safeguards include role-based access controls, secure audit trails, validated systems where applicable, reliable backup and recovery arrangements, and protection against unauthorized alteration. These controls help preserve the provenance of data and maintain a traceable history of its creation, processing, review, and modification.
Departures from approved protocols or established procedures must be documented, explained, and assessed for their potential impact on study conduct, data quality, and the interpretation of results. Deviations should be evaluated objectively rather than concealed because they complicate the findings. Unexpected observations, scientifically relevant negative results, and other findings that may affect the study’s conclusions must likewise be recorded and reported appropriately. Through protocol control, procedural standardization, equipment oversight, and comprehensive recordkeeping, GLP supports the reconstruction of a study from its underlying records and evidence. This reconstructability enables independent reviewers to assess the reliability of the reported findings and determine whether the conclusions are adequately supported by the experimental record.
Quality assurance, study reporting, archiving, and regulatory compliance
Quality assurance (QA), final reporting, and archival management provide the mechanisms through which compliance with GLP can be demonstrated beyond the immediate laboratory environment. Experimental findings acquire regulatory significance not merely because measurements have been obtained, but because the methods, observations, decisions, and conclusions are supported by a structured, traceable, and verifiable documentary record. Accordingly, GLP requirements extend across the entire study lifecycle, from initial planning and experimental conduct to final reporting and the long-term preservation of study-related materials.
The QA Unit conducts inspections in accordance with an established programme appropriate to the nature, scope, and progress of the studies. These inspections may focus on individual studies, specific experimental procedures, or general laboratory processes. QA personnel assess compliance by comparing actual practices with approved study protocols, applicable SOPs, and relevant GLP requirements. They verify whether study activities have been performed as planned, observations and measurements have been recorded accurately, deviations have been documented appropriately, and essential procedures have been conducted under suitable controls. The findings provide an independent assessment of the extent to which the study complies with established requirements.
QA findings are communicated to the Study Director and appropriate management personnel to facilitate the investigation, documentation, and correction of identified deficiencies. The final study report must present an accurate and comprehensive account of the study, including its objectives, methods, results, relevant observations, deviations, and conclusions. It should identify the personnel responsible for key study activities and provide sufficient information to enable an independent reader to understand how the findings were obtained and how the conclusions were derived.
The Study Director’s signed approval establishes accountability for the scientific content of the report and confirms that it accurately represents the study as conducted. Where required, the report must also include the applicable QA statement documenting the inspections performed and the dates on which findings were communicated to management and the Study Director. Archiving preserves the documentary and material evidence required for subsequent review, regulatory inspection, or scientific clarification.
Depending on the study and applicable requirements, archived materials may include the approved study protocol and amendments, raw data, the final report, relevant specimens and samples, retained test and reference items, and records of QA inspections. Archival systems must protect these materials against deterioration, unauthorized access, accidental destruction, and loss while ensuring their accessibility throughout the required retention period. Electronic records require appropriate safeguards, including access controls, backup arrangements, data integrity measures, continued readability, and reliable retrieval procedures.
Responsibilities for archival custody, retention, transfer, and authorized retrieval must be clearly defined to maintain the continuity and traceability of the study record. Regulatory authorities may inspect laboratory facilities and examine study documentation to assess compliance with applicable GLP requirements. Such inspections may evaluate organizational arrangements, the independence of the QA Unit, personnel qualifications and training, equipment management, data traceability, study documentation, and archival practices. Identified deficiencies may prompt requests for clarification or corrective action and, where the deficiencies compromise compliance or data reliability, may affect the regulatory acceptability of the study results.
The consequences depend on the nature and severity of the findings, the applicable regulatory framework, and their implications for the integrity and reliability of the data. GLP compliance should not be interpreted as a guarantee that every experiment is scientifically flawless or that every conclusion is correct. Rather, GLP provides a structured framework for ensuring that non-clinical studies are organized, conducted, documented, reported, and archived in a controlled and traceable manner. Scientific validity also depends on appropriate study design, suitable and reliable methods, and sound interpretation of the results.
Similarly, GLP compliance alone does not establish that a substance is safe for human use or environmental release; such determinations require the broader assessment of relevant scientific evidence and applicable regulatory criteria. Through independent quality oversight, accurate study reporting, secure archival management, and regulatory inspection, GLP establishes a verifiable link between experimental evidence and reported conclusions. These interconnected processes support the reconstruction and critical evaluation of study findings, strengthen regulatory accountability, and promote confidence in non-clinical data used to inform decisions concerning public health, animal welfare, and environmental protection.
Good manufacturing practice (GMP)
Good manufacturing practice (GMP) is a comprehensive regulatory and quality management framework designed to ensure that products intended for human or veterinary use are consistently manufactured, tested, packaged, stored, and distributed in accordance with established quality standards. It provides a systematic approach to manufacturing that ensures product quality, safety, identity, purity, strength, and reliability throughout the product lifecycle. GMP is particularly important in pharmaceutical manufacturing, biotechnology, vaccine production, medical device manufacturing, and other industries in which product deficiencies may adversely affect human health or animal welfare.
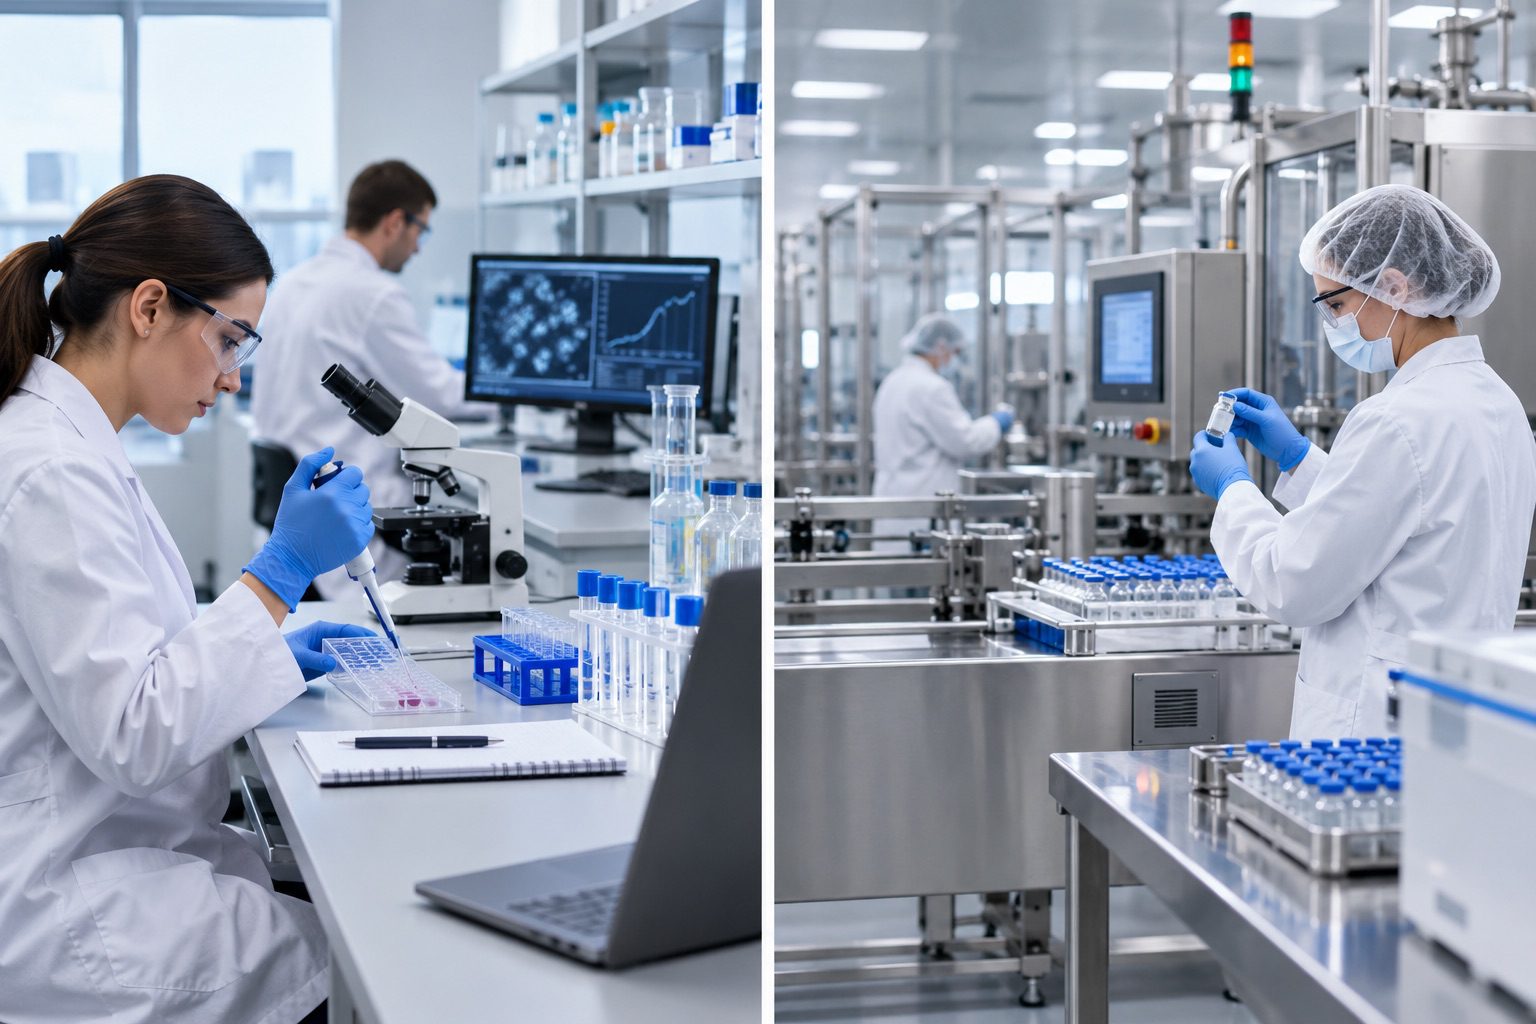
Its primary objective is to prevent manufacturing errors, contamination, cross-contamination, product deterioration, and uncontrolled process variations that could compromise the quality, safety, or efficacy of finished products. Unlike GLP, which primarily governs the conduct, documentation, and integrity of regulated nonclinical laboratory studies, GMP focuses on the control, consistency, and reproducibility of manufacturing processes. GLP ensures the reliability and traceability of data generated during regulated studies, whereas GMP ensures that manufactured products consistently meet predefined quality specifications. These complementary frameworks support different stages of the product lifecycle, linking the generation of reliable scientific evidence with the controlled production of products intended for commercial distribution or clinical use.
GMP implementation is governed by regulatory frameworks and guidelines established by national and international authorities. In the United States, the Food and Drug Administration (FDA) establishes current GMP requirements for pharmaceutical manufacturing under Title 21 of the Code of Federal Regulations (CFR), particularly Parts 210 and 211 for finished pharmaceuticals. In the European Union, GMP requirements are set out in the EU GMP guidelines, while the World Health Organization (WHO) provides internationally recognized GMP recommendations.
These frameworks address personnel qualifications and training, facility design and maintenance, equipment suitability, sanitation, production controls, laboratory testing, documentation, process validation, and quality assurance. Although specific requirements vary according to jurisdiction and product category, the central principle remains consistent: quality must be built into manufacturing processes rather than assessed solely through testing of the finished product. GMP emphasizes preventive quality assurance, operational consistency, and evidence-based decision-making.
Manufacturing activities must be performed in accordance with approved procedures, and critical operations must be documented to provide reliable evidence of compliance and traceability. Facilities must maintain appropriate environmental and hygienic conditions, personnel must receive role-specific training, and equipment must be appropriately qualified, calibrated, maintained, and operated. Raw materials, components, and packaging materials must be properly identified, handled, sampled, tested where required, and approved for use. In addition, manufacturing processes must be adequately controlled, deviations must be investigated, and corrective and preventive actions must be implemented when necessary to address identified deficiencies and reduce the risk of recurrence.
Through these integrated controls, GMP establishes a systematic framework for identifying, evaluating, and managing manufacturing risks. It protects patients, consumers, and animals by promoting consistent product quality, traceability, and conformity with applicable regulatory requirements and approved specifications. By integrating quality assurance into every relevant stage of production, GMP serves as a fundamental component of pharmaceutical quality management and responsible industrial manufacturing, supporting the consistent production of safe, effective, and high-quality products.
GMP organizational structure, personnel, and manufacturing controls
The organizational structure of GMP is designed to establish clear lines of responsibility, minimize conflicts of interest, and ensure independent oversight of manufacturing activities. The principal functional areas typically include production, quality control (QC), and QA. Although these functions must collaborate closely, their responsibilities should remain sufficiently independent to ensure that manufacturing operations are subject to objective quality assessments and appropriate oversight.
The production department is responsible for executing approved manufacturing instructions and ensuring that all operations comply with established procedures. Its activities typically include material dispensing, formulation, processing, equipment operation, packaging, and the completion of manufacturing records. Production personnel must verify material identities, adhere to specified process parameters, and promptly report abnormalities or unexpected events. Any departure from an approved instruction must be documented, investigated, and assessed through the established deviation-management system, with appropriate corrective actions implemented where necessary.
The QC department provides analytical and microbiological evidence to assess the conformity of materials and products with established specifications. QC laboratories examine incoming raw materials, packaging components, intermediate products, and finished batches using approved analytical methods. Depending on the product and its intended use, testing may include identity verification, assay determination, impurity profiling, dissolution testing, sterility testing, bacterial endotoxin analysis, and microbial enumeration. These results support decisions regarding material acceptance and product conformity. However, testing alone cannot ensure product quality or compensate for inadequately designed processes and uncontrolled manufacturing conditions. Quality must therefore be built into manufacturing processes and maintained through effective process controls.
The QA department oversees the implementation and effectiveness of the pharmaceutical quality management system. Its responsibilities commonly include document control and approval, deviation investigation, change control, corrective and preventive action (CAPA), internal audits, supplier qualification, training oversight, and the review of batch manufacturing records. QA also evaluates whether identified deficiencies represent isolated incidents or indicate broader systemic weaknesses. Through these activities, QA promotes continual improvement, supports regulatory compliance, and helps ensure that quality-related decisions are based on documented evidence and established procedures. Where required, QA or another appropriately authorized quality function is responsible for the final disposition and release of batches.
Personnel qualification and training are essential to effective GMP implementation. Employees must possess the appropriate qualifications, practical skills, and knowledge necessary to perform their assigned responsibilities. Training programs should cover relevant SOPs, personal hygiene, contamination prevention, data integrity, equipment operation, and applicable occupational safety requirements. Training effectiveness should be assessed where appropriate, and records must be maintained to demonstrate personnel competency. In controlled manufacturing environments, personnel must also comply with established gowning, hygiene, and access-control procedures to minimize contamination risks and protect product quality.
Manufacturing premises and equipment must be designed, operated, and maintained to prevent errors, contamination, and product mix-ups. Facility layouts should support orderly material and personnel flows, effective cleaning, appropriate maintenance, and the separation of incompatible operations where necessary. Depending on the product and manufacturing process, additional controls may include environmental monitoring, air filtration, pressure differentials, and the maintenance of specified temperature and humidity conditions. Equipment must be suitable for its intended purpose, maintained according to approved schedules, and calibrated when measurement accuracy is critical. Qualification and validation activities should provide documented evidence that facilities, equipment, and relevant processes consistently perform as intended.
Material management and control extend from supplier qualification and material receipt through identification, sampling, quarantine, testing, release, storage, and use in manufacturing. Materials must be handled and stored under appropriate conditions, with their status clearly identified throughout the supply chain and manufacturing process. Rejected, expired, or otherwise unsuitable materials must be appropriately labeled, segregated, and controlled to prevent their unintended use. Traceability records should enable materials to be linked to the relevant manufacturing operations and finished batches.
These organizational and operational controls establish accountability, strengthen independent quality oversight, and reduce the risk of preventable errors. By integrating competent personnel, clearly defined responsibilities, controlled facilities and equipment, effective material management, and a robust pharmaceutical quality management system, GMP provides the framework for consistently manufacturing products that meet predefined quality requirements and applicable regulatory standards.
GMP quality control, documentation, validation, and batch release
QC laboratories are integral to the GMP framework because their findings directly inform decisions regarding material acceptance, product quality, and batch release. Unlike exploratory laboratories, which primarily investigate hypotheses and generate new scientific knowledge, GMP laboratories apply validated or otherwise appropriately qualified analytical procedures to determine whether materials and products comply with predefined specifications. These activities must be performed using controlled procedures, suitable and qualified equipment, trained personnel, and documented acceptance criteria to ensure reliable, reproducible, and scientifically defensible results.
Typical QC activities include chromatographic assays, spectroscopic identification, potency determination, dissolution testing, impurity quantification, sterility testing, microbial-limit testing, and endotoxin analysis, depending on the product and its intended use. Before routine implementation, analytical procedures must be demonstrated to be suitable for their intended purpose through appropriate validation or verification, as applicable. Relevant performance characteristics may include accuracy, precision, specificity, linearity, range, detection limit, and quantitation limit. Instruments must be appropriately qualified and calibrated, while reference standards, reagents, and laboratory consumables must be controlled to ensure their suitability and traceability.
Laboratory investigations are essential when results fall outside established specifications or exhibit unexpected patterns or trends. An out-of-specification (OOS) result requires a documented, scientifically justified investigation rather than being disregarded or subjected to repeated testing without a valid rationale. Investigations should assess potential laboratory errors, analytical procedure performance, sample handling, equipment condition, and possible manufacturing-related causes. Where a failure is confirmed, appropriate actions may include batch rejection, an expanded investigation, corrective and preventive actions (CAPA), and regulatory notification, where required by applicable regulations. Such investigations help identify underlying causes, prevent recurrence, and protect product quality and patient safety.
Documentation provides the evidentiary foundation for demonstrating GMP compliance. Each batch must have sufficiently complete records to reconstruct its manufacturing, testing, and disposition history. These records typically include raw material batch identifiers, equipment used, processing parameters, personnel involved, in-process observations, packaging operations, deviations, and laboratory results. Documentation must be attributable, legible, contemporaneous, original or appropriately verified, and accurate, consistent with data integrity principles. Electronic records and computerized systems require appropriate access controls, audit trails, backup and recovery arrangements, and other safeguards commensurate with their intended use and applicable regulatory requirements. These controls help preserve the completeness, consistency, and reliability of data and prevent unauthorized alteration, deletion, or concealment of critical information.
Validation and qualification provide documented evidence that facilities, equipment, computerized systems, cleaning procedures, analytical methods, and manufacturing processes are suitable for their intended purposes. Equipment qualification commonly comprises Installation Qualification (IQ), Operational Qualification (OQ), and Performance Qualification (PQ), where applicable. Analytical method validation establishes that a procedure consistently performs as required for its intended application, while process validation provides evidence that a defined manufacturing process can reproducibly produce material meeting predetermined quality requirements. Cleaning validation demonstrates that residues and contaminants can be reduced to scientifically justified, acceptable limits. Following process validation, continued process verification and ongoing monitoring help establish whether process performance remains in a state of control during routine manufacturing.
Following manufacturing and testing, each batch undergoes a formal review of relevant documentation, analytical results, deviations, and other quality-related information before release. This review confirms that manufacturing was performed in accordance with approved procedures, applicable specifications were met, and any deviations or investigations were appropriately assessed and resolved. In the European Union, for example, an appropriately authorized Qualified Person (QP) must certify a medicinal product batch before its release for sale or supply, in accordance with applicable regulatory requirements. This process links laboratory evidence, manufacturing documentation, and regulatory accountability, ensuring that distribution decisions are supported by a comprehensive assessment of product quality and compliance with GMP requirements.
Regulatory compliance, risk management, and product lifecycle assurance
Regulatory compliance under GMP extends beyond the execution of individual manufacturing procedures. It requires an integrated pharmaceutical quality system capable of identifying hazards, controlling process variability, investigating failures, and maintaining consistent product quality throughout commercial manufacturing and the product lifecycle. Regulatory authorities assess whether manufacturers have effective systems in place to achieve these objectives and whether documented procedures accurately reflect actual operations.
GMP inspections may cover manufacturing facilities, laboratory operations, personnel training, equipment maintenance, cleaning procedures, computerized systems, material traceability, batch records, and quality management processes. Inspectors also evaluate deviation investigations, complaint handling, the effectiveness of corrective and preventive actions (CAPA), supplier oversight, and change control activities that may affect product quality. Inspection scope and frequency depend on the applicable regulatory framework, product category, manufacturing history, risk profile, and other relevant factors. Significant deficiencies may lead to regulatory observations, import restrictions, warning letters, product recalls, suspension of manufacturing activities, or other enforcement actions, depending on their nature and severity.
Quality risk management is fundamental to establishing and maintaining a scientifically justified control strategy. Manufacturers should proactively identify potential hazards, including contamination, product mix-ups, equipment malfunction, human error, and process instability, before they compromise product quality or patient safety. Risk assessment tools support the prioritization of controls based on factors such as the severity of potential harm, the probability of occurrence, and the detectability of failure. Within the pharmaceutical industry, ICH Q9, Quality Risk Management, provides principles and methodologies for systematic risk assessment and control, while ICH Q10, Pharmaceutical Quality System, establishes a framework for managing pharmaceutical quality throughout the product lifecycle. Risk management activities should be proportionate to the level of risk, supported by scientific evidence, and reviewed when new information becomes available.
The International Council for Harmonisation of Technical Requirements for Pharmaceuticals for Human Use (ICH) develops harmonized guidelines to support the quality, safety, and efficacy of medicinal products. In particular, ICH Q10 describes a comprehensive pharmaceutical quality system spanning pharmaceutical development, technology transfer, commercial manufacturing, and product discontinuation. It emphasizes management responsibility, process performance and product quality monitoring, CAPA, change management, and continual improvement. Together with applicable GMP requirements, these principles support consistent product quality and effective oversight across the product lifecycle.
Deviation management and CAPA provide structured mechanisms for addressing unexpected events, determining their underlying causes, and preventing recurrence. Deviations may arise from equipment malfunctions, environmental excursions, documentation discrepancies, out-of-specification results, or departures from approved procedures. Investigations should establish the most probable root cause based on objective evidence, assess the potential impact on affected batches, and determine whether other products, processes, or systems may be affected. Corrective actions address identified problems and their causes, whereas preventive measures seek to reduce the likelihood of recurrence or comparable failures. CAPA effectiveness checks are essential to verify that implemented measures achieve their intended outcomes and that the identified risks have been adequately controlled.
Change control is equally important because modifications to equipment, raw materials, suppliers, analytical methods, computerized systems, facilities, or manufacturing parameters may affect product quality, process performance, or regulatory compliance. Proposed changes should undergo a documented assessment of their potential impact on product quality, patient safety, validated states, and regulatory commitments. Changes must be reviewed and approved by authorized personnel before implementation, with appropriate qualification, validation, verification, and training performed as necessary. Where changes affect approved product information or regulatory commitments, the relevant notification or approval requirements must also be evaluated and fulfilled before implementation, as applicable.
GMP responsibilities continue beyond batch release. Manufacturers must maintain appropriate storage and distribution controls, investigate product quality complaints, monitor emerging quality signals, and implement recall procedures when warranted. Temperature-sensitive products may require validated storage conditions, qualified transportation systems, and continuous or appropriately defined temperature monitoring to preserve product quality throughout distribution. Stability studies provide evidence that products remain within approved specifications under defined storage conditions throughout their assigned shelf life. Post-market information, including complaints, stability trends, and other relevant quality data, should feed back into the pharmaceutical quality system to support ongoing risk assessment and continual improvement.
GMP establishes a lifecycle-oriented framework in which product quality is maintained through preventive controls, documented and traceable evidence, effective oversight, and continual evaluation. Its effectiveness depends not merely on the existence of written procedures but on their consistent implementation, scientifically sound investigations, reliable data, and clear organizational accountability. By integrating regulatory compliance, quality risk management, deviation handling, CAPA, change control, and post-market quality oversight, manufacturers can strengthen process reliability, maintain regulatory compliance, and support the consistent delivery of safe, effective, and high-quality medicinal products to patients.
Regulatory foundations and operational distinctions between GLP and GMP
The distinction between GLP and GMP reflects two complementary regulatory frameworks governing pharmaceutical development and production. GLP focuses on the reliability, integrity, and traceability of non-clinical study data, whereas GMP ensures the consistent manufacture of products that meet predefined quality standards. These frameworks establish distinct compliance requirements that influence laboratory and manufacturing operations, organizational responsibilities, documentation practices, and quality oversight throughout the pharmaceutical development lifecycle.
Table 1 summarizes the principal regulatory and operational differences between GLP and GMP, highlighting their respective roles in pharmaceutical quality assurance. Under GLP, key requirements include adherence to approved study protocols, the use of scientifically appropriate and fit-for-purpose analytical methods, control of experimental conditions, preservation of raw data, and comprehensive documentation to enable the reconstruction, verification, and independent assessment of non-clinical safety studies.
Table 1. Differences between GLP and GMP laboratory testing
| Topic | Good Laboratory Practice (GLP) | Good Manufacturing Practice (GMP) |
| Study Director | Single appointed individual responsible for overall conduct, scientific integrity, and compliance of the study. Designated by Testing Facility Management. | No equivalent role. Responsibility is distributed across production, QC, and QA systems; no single study-level authority exists. |
| Quality Assurance / Quality Control | Independent Quality Assurance Unit (QAU) audits study phases, verifies compliance, and reports directly to management. Does not perform testing or control work. | Quality Control (QC) unit performs testing and has authority to accept or reject materials/products against specifications; QA oversees system compliance and release processes. |
| Testing Facility Management | Ensures qualified Study Director is assigned, maintains independent QAU, and provides adequate resources, equipment, and trained personnel for studies. | Ensures trained supervisors, validated systems, and compliance with SOPs; responsible for maintaining controlled manufacturing and quality systems. |
| Type of Testing Conducted | Non-clinical, open-ended experimental studies for regulatory safety assessment (e.g., toxicology, pharmacokinetics, environmental studies). | Routine, specification-driven testing of raw materials, in-process samples, and finished products to confirm compliance for release. |
| Facility Requirements | Facilities must support study integrity, minimize contamination/mix-ups, and ensure traceability; infrastructure requirements less prescriptive. | Highly regulated facility design with strict environmental controls, segregation of areas, controlled utilities (HVAC, water systems, cleaning zones). HVAC-Heating, Ventilation, and Air Conditioning systems |
| Equipment | Must be suitable for study purpose, calibrated, and fully documented to ensure study reconstructability; focus on experimental accuracy. | Must be qualified (IQ/OQ/PQ where applicable), validated for manufacturing use, and routinely calibrated to ensure consistent product quality. IQ-installation qualificationOQ-operational qualificationPQ-performance qualification |
| Standard Operating Procedures (SOPs) | Developed by qualified personnel and approved by management; may be adapted per study requirements. | Strictly controlled, formally approved by QA/QC; used consistently with minimal deviation across production activities. |
| Study Protocol / Process Control | Each study requires a detailed protocol defining objectives, methods, and analysis; deviations require documentation and justification. | No study protocols; manufacturing follows validated processes and batch records rather than experimental protocols. |
| Master Schedule / Indexing | QAU maintains a master schedule listing all ongoing and completed studies for audit tracking. | Not formally required as a regulatory element in GMP systems. |
| Records and Reports | Full raw data traceability required; all entries signed, dated, and attributable. Archiving required for long-term regulatory access (typically 2–5+ years depending on use). | Batch records require dual verification (operator + reviewer). Records retained typically until product expiry plus defined period; focus on batch traceability. |
| CAPA System | Not a mandatory structural requirement (may exist but not central to GLP framework). | Mandatory Corrective and Preventive Action (CAPA) system to investigate deviations, prevent recurrence, and maintain continuous quality improvement. |
By contrast, GMP focuses on the control and consistency of manufacturing processes, traceability of materials, qualification of equipment, personnel training and competency, deviation investigation, and documented quality control testing. Together, these requirements help ensure that non-clinical study findings are reliable and that manufactured products consistently conform to established quality specifications.
These distinctions are reflected in organizational responsibilities, facility controls, record-management systems, and the implementation of quality assurance activities. Despite their differences in regulatory purpose and operational scope, GLP and GMP share a common foundation of predefined procedures, comprehensive documentation, and systematic oversight. Their complementary application helps ensure the generation of scientifically reliable safety data and the consistent manufacture of pharmaceutical products that meet applicable regulatory standards and quality requirements.
References
Bedson, P., & Sargent, M. (1996). The development and application of guidance on equipment qualification of analytical instruments. Accreditation and Quality Assurance, 1, 265–274.
Bridges, D. (2005). Compliance requirements for equipment and instrumentation in GLP studies. Society of Quality Assurance, 21st SQA Annual Meeting Preconference Training, First Global QA Conference, 1–17.
Brockmeyer, P. (2006). Process validation fundamentals. 29th Biological GMP and Validation Education, 21–28.
Cho, K. H., Kim, J. S., Jeon, M. S., Lee, K., Chung, M. K., & Song, C. W. (2009). Basic principles of the validation for Good Laboratory Practice institutes. Toxicological Research, 25(1), 1–8.
European Union. (2003). Guide to good manufacturing practice: Annex 15.
Haider, S. I. (2002). Validation standard operating procedures: A step-by-step guide for achieving compliance in the pharmaceutical, medical device, and biotech industries. St. Lucie Press.
Huber, L. (2005). Compliance requirements for equipment and instrumentation in GLP studies in gap analysis, risk assessment, and master validation plan.
Jarvis, B. (2014). Good manufacturing practice. In C. A. Batt & M. L. Tortorello (Eds.), Encyclopedia of food microbiology (2nd ed., pp. 106–115). Academic Press.
Thomas, B. F., & ElSohly, M. A. (2016). Quality control and stability assessment. In The analytical chemistry of cannabis (pp. 83–99). Elsevier.
World Health Organization. (n.d.). Good manufacturing practices for biological products. https://www.who.int/biologicals/vaccines/good_manufacturing_practice/en/
MicroChem Laboratory. (n.d.). What is the difference between GLP and GMP laboratory testing regulations?https://www.microchemlab.com/information/what-difference-between-glp-and-gmp-laboratory-testing-regulations
Discover more from Microbiology Class
Subscribe to get the latest posts sent to your email.