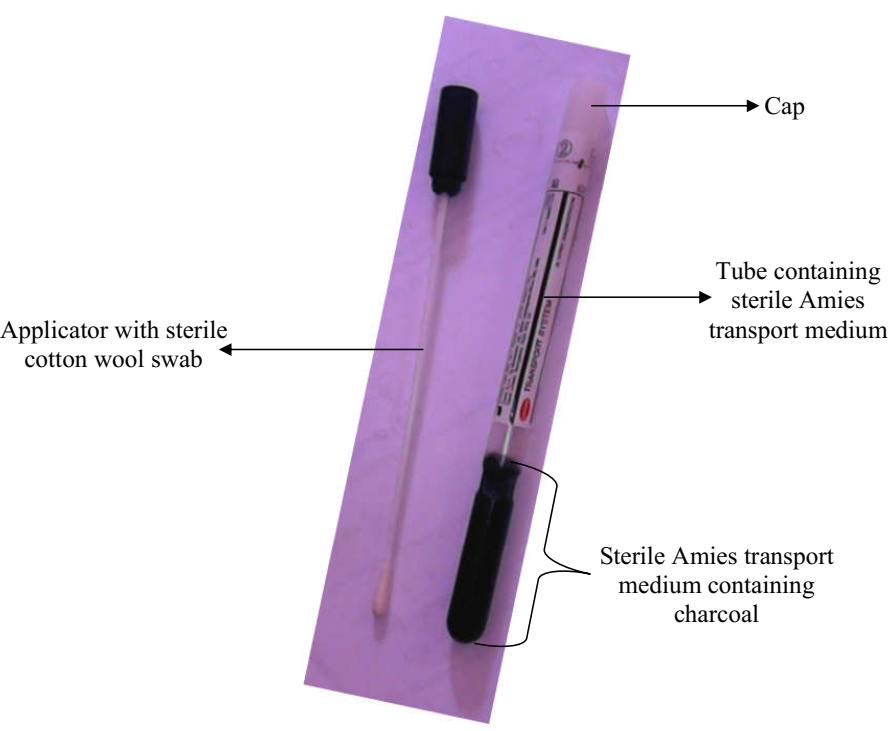
Good laboratory practice (GLP) is simply defined as a laboratory quality measure or protocol that is concerned with the organizational process and the conditions under which laboratory experimentations (inclusive of other non-clinical studies) are planned, performed, monitored, recorded, archived, retrieved and reported. GLP is a structured system of management controls that governs how laboratory studies are planned, performed, monitored, recorded, reported, and archived. It is primarily applied in non-clinical research such as toxicology, environmental safety, and pharmaceutical testing to ensure that generated data are reliable, reproducible, and acceptable to regulatory authorities. At its core, GLP emphasizes standardization and traceability. This involves the use of validated methods, calibrated equipment, and well-defined Standard Operating Procedures (SOPs). Each study must follow a pre-approved protocol, and any deviations are documented and justified. Personnel roles and responsibilities are clearly assigned, and staff must be appropriately trained and qualified.
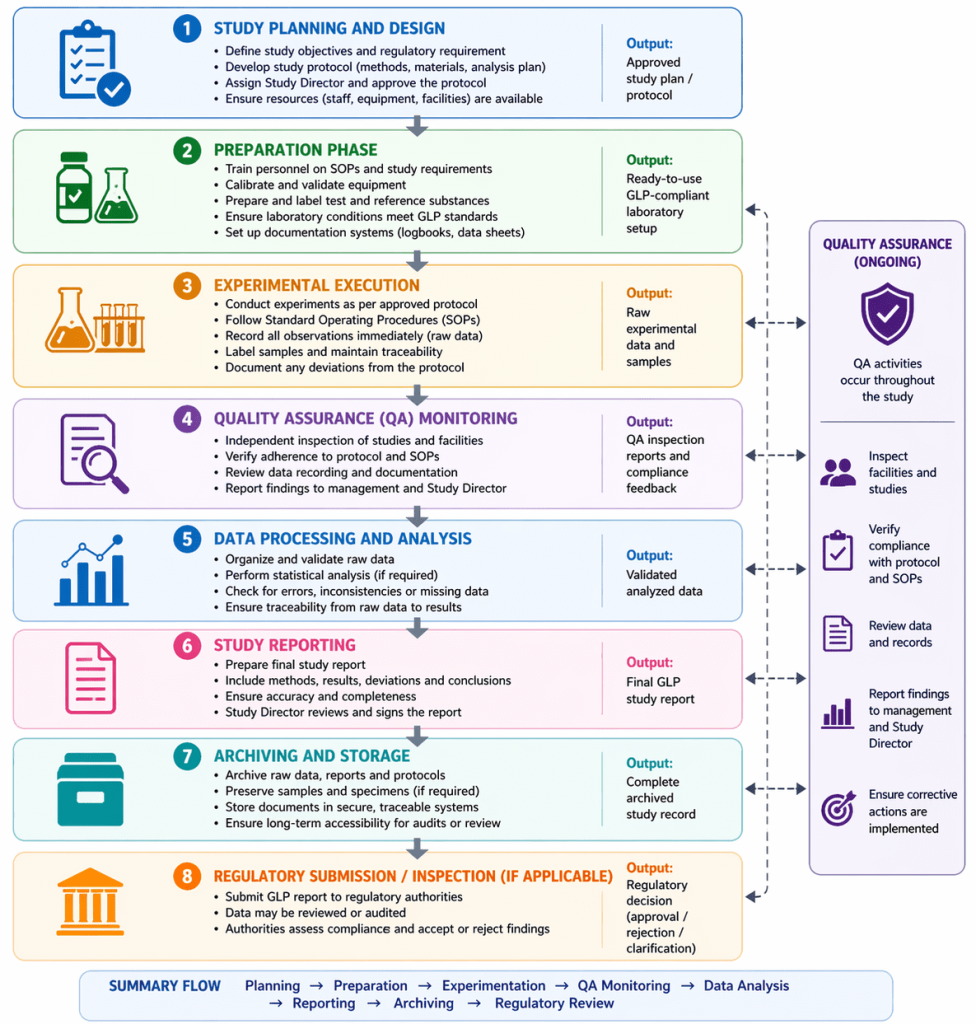
A key feature of GLP is rigorous documentation. All raw data, observations, and results are recorded promptly and accurately, creating a transparent audit trail. Quality assurance (QA) units operate independently to inspect procedures and verify compliance throughout the study lifecycle. GLP also covers proper handling of test and reference substances, maintenance of facilities, and secure archiving of records and samples. By enforcing consistency and accountability, GLP minimizes errors, reduces bias, and enhances confidence in scientific findings, especially when used to support regulatory decisions. Good Laboratory Practice is a cornerstone of modern scientific research, particularly in regulated environments. By enforcing standardized procedures, rigorous documentation, and independent quality assurance, GLP ensures that laboratory data are credible, reproducible, and suitable for regulatory decision-making. In an era where scientific findings directly impact public health, environmental protection, and technological advancement, adherence to GLP is not optional; it is essential. A simplified workflow of GLP is shown in Figure 1.
Objectives and Importance of Good Laboratory Practice (GLP)
The primary objective of Good Laboratory Practice (GLP) is to ensure the generation of high-quality, reliable, and reproducible scientific data. This is particularly critical in non-clinical research, where study outcomes inform regulatory decisions such as the approval of pharmaceuticals, agrochemicals, and industrial chemicals. GLP establishes a standardized framework that minimizes variability, reduces errors, and enhances confidence in laboratory findings across different institutions and jurisdictions.
Data Integrity
Data integrity refers to the accuracy, completeness, and consistency of data throughout its lifecycle. Under GLP, all data must be recorded promptly, clearly, and in a manner that prevents unauthorized alteration. This includes maintaining raw data in its original form and ensuring that any modifications are fully traceable through an audit trail. Practices such as proper documentation, controlled access to records, and secure data storage systems are essential components. Ensuring data integrity prevents scientific misconduct, such as fabrication or falsification, and guarantees that conclusions drawn from the data are based on authentic and verifiable evidence.
Reproducibility
Reproducibility is a cornerstone of scientific validity. GLP enforces the use of standardized methods, validated procedures, and controlled experimental conditions to ensure that studies can be repeated with consistent results. This includes detailed documentation of methodologies, calibration of equipment, and adherence to Standard Operating Procedures (SOPs). When experiments are reproducible, independent laboratories can confirm findings, strengthening the credibility of the results. Lack of reproducibility can undermine scientific trust and lead to rejection of data by regulatory authorities.
Accountability
Accountability within GLP ensures that every aspect of a study is traceable to specific individuals and actions. Roles and responsibilities are clearly defined, from the Study Director to technical staff, and all activities are documented. This traceability allows for effective oversight and facilitates investigations in case of discrepancies or errors. By assigning responsibility and maintaining transparent records, GLP promotes ethical conduct and ensures that all personnel adhere to established standards.
Regulatory Acceptance
One of the most significant advantages of GLP compliance is the acceptance of data by regulatory authorities worldwide. Organizations such as environmental and health agencies require that submitted data adhere to GLP standards before they are considered for risk assessment or product approval. GLP harmonization across countries reduces duplication of studies, saving time and resources. Without GLP compliance, data may be deemed unreliable, leading to delays, increased costs, or even rejection of applications, which can ultimately impact public health and environmental safety.
Organizational Structure and Personnel Responsibilities
A clearly defined organizational structure is fundamental to compliance with Good Laboratory Practice (GLP). It ensures that responsibilities are unambiguous, workflows are controlled, and accountability is maintained throughout the lifecycle of a study. Each role within the structure contributes to the integrity, traceability, and reliability of generated data.
Test Facility Management
Test facility management holds ultimate administrative responsibility for GLP compliance. Their role extends beyond oversight to actively establishing the infrastructure necessary for high-quality scientific work. This includes allocating sufficient resources both human and material such as qualified personnel, validated equipment, and appropriate laboratory space. Without adequate resourcing, even well-designed studies may fail to meet GLP standards. Management is also responsible for the development, approval, and enforcement of Standard Operating Procedures (SOPs), which standardize routine operations and minimize variability. Equally critical is the implementation of structured training programs to ensure that all staff possess the technical competence required for their roles. Training must be continuous and documented. Another core responsibility is establishing and maintaining an independent Quality Assurance (QA) unit. Management must ensure that this unit has the authority, resources, and autonomy to perform audits and inspections effectively. In essence, management sets the tone for a culture of compliance and scientific integrity.
Study Director
The Study Director is the central scientific authority for an individual study and serves as the single point of control. This role is pivotal because it integrates scientific design with regulatory compliance. The Study Director is responsible for drafting the study protocol, which defines the objectives, methodology, and acceptance criteria. During the experimental phase, the Study Director oversees execution to ensure adherence to the protocol and GLP requirements. This includes managing deviations, coordinating personnel, and ensuring that data collection is accurate and complete. Importantly, the Study Director must interpret the results objectively and compile the final report. Accountability is a defining feature of this role. The Study Director bears ultimate responsibility for the validity, integrity, and credibility of the study, meaning that any deficiencies in design, execution, or reporting ultimately fall under their remit.
Quality Assurance (QA) Unit
The Quality Assurance unit functions as an independent monitoring body within the GLP framework. Its independence from study personnel is critical to ensuring objective evaluation. The QA unit systematically inspects ongoing studies to verify that procedures are being conducted in accordance with approved protocols and SOPs.In addition to real-time inspections, QA personnel audit raw data, records, and final reports to ensure completeness, consistency, and accuracy. They also assess whether any deviations have been properly documented and justified. Their findings are reported directly to management, not to the Study Director, preserving impartiality. This continuous oversight acts as a control mechanism to detect errors, prevent non-compliance, and maintain confidence in the generated data.
Study Personnel
Study personnel include all technical staff involved in conducting experiments and handling data. Their competence is essential for the practical implementation of GLP principles. Each individual must be adequately trained, both theoretically and practically, for their assigned tasks. Personnel must have a clear understanding of their responsibilities and strictly adhere to SOPs and study protocols. Even minor deviations at this level can compromise data integrity. Therefore, consistent performance and procedural discipline are critical. Training records must be meticulously maintained to demonstrate competency and compliance during audits or inspections. In practice, study personnel form the operational backbone of GLP, translating protocols and procedures into reliable experimental outcomes.
Standard Operating Procedures (SOPs)
Standard Operating Procedures (SOPs) are formally documented, step-by-step instructions that describe how routine laboratory tasks must be executed under controlled conditions. Within a Good Laboratory Practice (GLP) framework, SOPs function as the operational backbone of consistency, ensuring that experimental procedures are performed in a standardized and reproducible manner. By eliminating ambiguity in how tasks are carried out, SOPs minimize procedural variability and strengthen the reliability of generated data. A critical feature of SOPs is that they must be clearly written and readily accessible to all relevant personnel. This means using precise, unambiguous language, avoiding unnecessary jargon where possible, and structuring instructions logically (e.g., sequential steps, safety notes, required materials). Accessibility also implies that SOPs are stored in centralized systems either physical manuals or digital repositories so staff can easily consult them during routine operations.
SOPs must also be regularly reviewed and updated to remain scientifically and technically relevant. Laboratory methods, instrumentation, and regulatory requirements evolve over time; therefore, periodic revision ensures that SOPs reflect current best practices. Version control is essential, with each update documented, approved, and communicated to staff to prevent the use of obsolete procedures. Another key aspect is that SOPs cover all critical laboratory processes, including equipment operation, calibration, maintenance, sample collection and handling, data recording, and safety procedures. Comprehensive coverage ensures that no essential step is left to individual interpretation, reducing the risk of inconsistency. SOPs play a central role in reducing human error, enhancing traceability, and ensuring that procedures are performed uniformly regardless of the operator. This standardization is crucial for producing high-quality, reproducible data that meet regulatory and scientific standards.
Study Planning and Protocol Development
Within GLP, study planning and protocol development represent a critical foundational phase that determines the scientific validity, regulatory acceptability, and operational efficiency of a study. A GLP-compliant study must begin with a formally approved protocol, which serves as the authoritative document guiding all experimental activities. It ensures that the study is conducted in a structured, standardized, and reproducible manner, minimizing ambiguity and reducing the risk of errors or bias.
A well-developed protocol is not simply a procedural outline; it is a legally and scientifically binding document that defines the scope, objectives, and execution strategy of the study. It must be reviewed and approved by the Study Director and, where applicable, test facility management before the study begins. The protocol also acts as a reference point for auditors, inspectors, and regulatory authorities evaluating the study. Study planning and protocol development under GLP provide the structural and scientific framework for conducting reliable and reproducible research. By clearly defining every aspect of the study in advance and maintaining strict control over any changes, GLP ensures that the resulting data are robust, transparent, and suitable for regulatory evaluation.
Components of a study protocol include:
Study title and objectives: The study title should be concise yet descriptive, accurately reflecting the purpose and nature of the investigation. It enables clear identification of the study in records and archives. The objectives must be explicitly stated, outlining what the study intends to achieve. These may include determining toxicity levels, assessing environmental impact, or evaluating the efficacy of a compound. Clearly defined objectives are essential because they guide the experimental design and ensure that the study remains focused and relevant. Ambiguous objectives can lead to misinterpretation of results or poorly designed experiments.
Description of test and control substances: A detailed description of all materials used in the study is essential for reproducibility and data integrity. This includes the test substance (e.g., a chemical or pharmaceutical compound) and any control or reference substances. Information should cover identity, chemical composition, purity, concentration, stability, and storage conditions. Proper characterization ensures that results can be accurately attributed to the substance being tested. Control substances, such as negative or positive controls, provide a baseline for comparison and help validate the experimental system.
Experimental design and methodology
The experimental design is the backbone of the protocol, detailing how the study will be conducted. It includes the selection of test systems (e.g., animal models, cell cultures), group allocation, sample size, dosing regimens, and duration of the study. Methodology must be described in sufficient detail to allow replication by another laboratory. This includes references to SOPs and validated methods. A robust design minimizes variability, controls confounding factors, and ensures that the data generated are scientifically sound and defensible.
Statistical methods: Statistical planning is essential to ensure that the study generates meaningful and interpretable data. The protocol must specify the statistical tests to be used, the rationale for their selection, and the criteria for significance. This includes defining sample sizes based on power calculations, identifying endpoints, and outlining how data will be analyzed. Proper statistical design prevents underpowered studies and reduces the risk of false conclusions. It also enhances the credibility of the results in regulatory submissions.
Criteria for data interpretation: The protocol must define how results will be evaluated and interpreted. This includes establishing thresholds, acceptance criteria, and decision rules. For example, it may specify what constitutes a significant toxic effect or a meaningful biological response. Clear interpretation criteria ensure consistency and objectivity, reducing the influence of subjective judgment. They also facilitate regulatory review by providing a transparent framework for assessing outcomes.
Protocol amendments and deviations: During the course of a study, unforeseen circumstances may necessitate changes to the original protocol. Under GLP, any such changes must be formally documented as protocol amendments. These amendments must be justified, approved by the Study Director, and communicated to all relevant personnel. Additionally, any unplanned deviations from the protocol must be recorded, explained, and assessed for their impact on the study’s integrity. This rigorous documentation ensures full traceability and maintains the credibility of the study.
Equipment and Instrumentation in GLP
Reliable equipment and instrumentation form the backbone of any laboratory operating under GLP. The accuracy, precision, and reproducibility of experimental data depend heavily on the proper functioning of instruments. Therefore, GLP establishes strict requirements to ensure that all equipment used in a study consistently performs within acceptable limits.
Reliable equipment is essential for generating accurate data. Therefore, GLP requires:
- Regular calibration and maintenance of equipment
- Proper installation and validation of equipment
- Documentation of performance checks on equipment
Regular calibration and maintenance: These steps are fundamental to ensuring measurement accuracy. Calibration involves comparing an instrument’s readings against a known standard to verify its correctness. This must be performed at defined intervals based on manufacturer recommendations, frequency of use, and criticality of the measurement. Maintenance, on the other hand, includes routine servicing such as cleaning, lubrication, replacement of worn parts, and software updates. Preventive maintenance minimizes the risk of unexpected breakdowns and ensures consistent performance over time. Failure to calibrate or maintain equipment can introduce systematic errors, compromising the validity of study results.
Proper installation and validation: Installation Qualification (IQ) ensures that the equipment is installed correctly according to manufacturer specifications and environmental requirements (e.g., temperature, humidity, electrical supply). Operational Qualification (OQ) verifies that the instrument functions as intended across its operating range, while Performance Qualification (PQ) confirms that it performs reliably under actual working conditions. Together, these validation steps ensure that the equipment is fit for its intended purpose before being used in any GLP study.
Documentation of performance checks: Routine performance checks such as running control samples, system suitability tests, or diagnostic routines must be conducted and recorded. These checks help detect deviations or malfunctions early, allowing corrective actions to be taken before data integrity is affected. Each instrument must also have a dedicated logbook, either in physical or electronic form. This logbook records details such as usage history, calibration dates, maintenance activities, repairs, and any observed issues or deviations. By maintaining a complete and traceable record, laboratories can demonstrate that all data generated are linked to properly functioning, validated equipment, thereby ensuring compliance with GLP standards and reinforcing confidence in the results.
Test and Reference Substances
GLP places strong emphasis on the proper management of test and reference substances because these materials form the foundation of any experimental study. Test substances are the chemicals, drugs, biological agents, or environmental compounds being investigated, while reference substances serve as standards or controls used for comparison. The reliability of experimental outcomes depends heavily on how well these substances are characterized, handled, and documented.
Requirements for test and reference substances include:
- Proper labeling with identity, concentration, and storage conditions
- Documentation of source and purity
- Stability testing where necessary
Proper labeling and identification
Every test and reference substance must be clearly and permanently labeled. Labels typically include the identity of the substance, batch or lot number, concentration or potency (where applicable), storage requirements, and expiry date. This ensures that there is no ambiguity during experimental procedures. Mislabeling or unclear identification can lead to serious errors, including incorrect dosing, invalid results, or irreproducible findings. In GLP-compliant laboratories, labeling is standardized and traceable throughout the entire study lifecycle.
Documentation of source and purity
GLP requires full traceability of substances back to their source. This includes documenting the manufacturer or supplier, procurement date, batch information, and certificate of analysis. Purity is especially important because contaminants or impurities can significantly alter biological or chemical responses in experiments. For example, a pharmaceutical compound with unknown impurities may produce misleading toxicity or efficacy results. Therefore, laboratories must maintain verified records confirming the chemical composition and quality of each substance used.
Stability testing and storage conditions
Many substances degrade over time due to exposure to light, heat, moisture, or oxygen. GLP therefore requires stability testing to determine how long a substance remains chemically and biologically stable under specified conditions. This information guides storage protocols, such as refrigeration, freezing, or protection from light. Proper storage ensures that substances maintain their intended properties throughout the study duration. Failure to control stability can introduce variability and compromise the validity of results.
Importance of test and reference substances for reproducibility and reliability
Accurate characterization and handling of test and reference substances ensure that experimental findings are both meaningful and reproducible. When substances are well-defined and consistently managed, other researchers can replicate the study under similar conditions and obtain comparable results. This is essential for regulatory acceptance and scientific credibility, particularly in fields like drug development and toxicology, where decisions directly affect human health and environmental safety.
Facilities and Environmental Conditions
The physical environment in which laboratory studies are conducted is another critical component of GLP. Proper facility design and environmental control are essential for preventing contamination, ensuring safety, and maintaining the integrity of experimental data. Even minor environmental inconsistencies can significantly affect sensitive biological or chemical experiments.
Key considerations for facilities and environmental conditions include:
- Separation of different activities (e.g., sample preparation, analysis)
- Controlled environmental conditions (temperature, humidity, light)
- Adequate storage areas for samples and reagents
Separation of laboratory activities
GLP requires that different laboratory activities be physically separated to avoid cross-contamination and operational interference. For example, sample preparation, chemical analysis, microbiological testing, and storage of reagents should occur in distinct designated areas. This separation prevents accidental mixing of substances, reduces contamination risk, and ensures that each stage of an experiment is conducted under controlled conditions. In high-standard laboratories, even directional workflow is designed to minimize cross-contact between “clean” and “dirty” zones.
Controlled environmental conditions
Environmental parameters such as temperature, humidity, airflow, and light exposure must be carefully controlled and continuously monitored. Many chemical reactions and biological systems are highly sensitive to environmental fluctuations. For instance, enzyme activity may decrease at inappropriate temperatures, or volatile compounds may degrade under high humidity. GLP-compliant laboratories often use environmental monitoring systems that continuously record conditions and trigger alarms when deviations occur. This ensures consistency across experiments and reduces variability caused by external factors.
Cleanliness and contamination control
Maintaining a clean laboratory environment is essential for preventing contamination of samples and reagents. Regular cleaning schedules, sterilization procedures, and use of protective equipment such as laminar flow hoods and biosafety cabinets are standard practices. Waste disposal systems must also be properly designed to prevent cross-contamination and environmental hazards. Contamination control is especially important in microbiology, toxicology, and pharmaceutical testing, where even trace contaminants can distort results.
Adequate storage facilities
Proper storage of samples, reagents, and reference materials is another essential requirement under GLP. Storage areas must be secure, organized, and environmentally controlled to maintain the stability of stored materials. This includes refrigerators, freezers, and chemical storage cabinets designed for specific types of substances. Labels and inventory systems ensure traceability and prevent mix-ups or accidental misuse. In addition, access to storage areas is often restricted to authorized personnel only, enhancing security and accountability.
Impact of facilities and environmental conditions on data integrity and reproducibility
Well-maintained facilities and controlled environmental conditions directly contribute to the generation of reliable and consistent data. When environmental variables are minimized and laboratory spaces are properly organized, experimental variability is reduced, allowing for more accurate interpretation of results. This is particularly important in regulatory science, where data must withstand rigorous scrutiny and often be reproducible across different laboratories and countries.
Data Recording and Documentation
Data recording and documentation are among the most fundamental pillars of GLP, as they ensure that all scientific findings are transparent, traceable, and verifiable. Without accurate documentation, even well-designed experiments lose scientific and regulatory value. GLP therefore requires that all data generated during a study be recorded systematically, promptly, and in a manner that preserves its integrity.
All observations and measurements must be recorded as soon as they are obtained to avoid memory-based errors or selective reporting. Records must be legible, clear, and permanent, ensuring that they can be reviewed and interpreted accurately by others, including auditors and regulatory authorities. In addition, every piece of recorded data must be attributable, meaning it must clearly identify the individual who generated or recorded it. This is typically achieved through signatures, initials, or secure electronic identification systems. Secure storage of data is also essential. Both paper-based and electronic records must be protected against loss, damage, unauthorized access, or tampering. Laboratories are required to implement controlled access systems, backup procedures, and archiving protocols to ensure long-term preservation of data.
Raw Data
Raw data refer to the original records of observations and results generated during a study. These may include handwritten notes, instrument printouts, digital files, photographs, or automatically recorded outputs from laboratory equipment. A key GLP requirement is that raw data must be preserved exactly as they were originally recorded, without alteration or deletion. If electronic systems are used, they must be validated to ensure data integrity and prevent unauthorized modifications.
Audit Trail
The audit trail is a critical mechanism for ensuring transparency and accountability in data management. Any modification made to recorded data must be fully documented. This includes:
- The specific reason for the change
- The identity of the individual making the modification
- The exact date and time of the change
Original data must remain visible even after corrections are made; they should not be erased or overwritten. Instead, corrections are typically made in a way that allows both the original entry and the updated information to be reviewed.
Quality Assurance and Inspections
The Quality Assurance (QA) unit is a fundamental component of GLP compliance, ensuring that all laboratory activities are conducted in accordance with established protocols, regulatory requirements, and internal SOPs. Its primary role is to provide an independent verification system that monitors study conduct, data integrity, and overall laboratory performance. Unlike study personnel, QA operates independently from experimental activities, which allows for objective assessment and unbiased reporting. QA functions as the “internal regulator” of the laboratory, continuously evaluating whether scientific procedures are being followed correctly and whether the generated data are reliable and defensible. This is essential in regulated environments where laboratory findings may influence critical decisions in pharmaceuticals, environmental safety, or chemical risk assessment.
Types of Inspections
QA inspections are structured into three major categories, each serving a distinct purpose in maintaining GLP compliance:
1. Study-based inspections
Study-based inspections focus on individual ongoing or completed studies. QA personnel verify that experiments are conducted strictly according to the approved study protocol. This includes checking raw data entries, confirming correct use of test substances, ensuring adherence to timelines, and verifying that deviations are properly documented and justified. These inspections are often conducted at multiple stages of a study to ensure continuous compliance rather than end-point validation only.
2. Facility inspections
Facility inspections evaluate the overall laboratory infrastructure and working environment. This includes assessment of equipment maintenance, cleanliness, environmental controls (such as temperature, humidity, and contamination prevention), storage conditions for samples and reagents, and adequacy of safety measures. The goal is to ensure that the physical environment does not compromise data quality or personnel safety.
3. Process inspections
Process inspections examine specific laboratory procedures or workflows that are routinely used across multiple studies. These may include sample preparation techniques, analytical methods, calibration procedures, or data recording systems. The objective is to ensure that these processes are standardized, validated, and consistently applied by all personnel.
Reporting and Corrective Actions
After each inspection, the QA unit prepares detailed reports documenting observations, deviations, and areas of non-compliance. These reports are submitted directly to laboratory management and the Study Director. Importantly, QA does not implement changes directly but ensures that corrective and preventive actions (CAPA) are developed and executed by responsible personnel.
Continuous Monitoring and Compliance Improvement
QA is not a one-time checkpoint but a continuous oversight mechanism embedded throughout the laboratory lifecycle. By systematically identifying errors, procedural weaknesses, and compliance gaps, QA helps improve laboratory performance over time. This ongoing monitoring strengthens data credibility, reduces the risk of regulatory rejection, and ensures that all scientific outputs meet internationally accepted GLP standards.
Reporting of Results
At the conclusion of a GLP study, the preparation of a comprehensive final report is a critical step that consolidates all experimental activities, observations, and outcomes into a single authoritative document. The final report serves as the primary evidence of what was performed during the study and is essential for regulatory review, scientific validation, and potential future audits. The report must accurately reflect the study findings without distortion, omission, or selective presentation of data. This includes both positive and negative results, as well as any unexpected observations that occurred during the study. Complete transparency is required to ensure the scientific integrity of the work. All relevant data and observations generated during the study must be included or properly referenced within the report, ensuring traceability back to the original raw data. A fundamental requirement of GLP is that the final report must be formally signed and dated by the Study Director. This signature confirms that the Study Director accepts full responsibility for the accuracy, completeness, and compliance of the report with the approved study protocol and GLP principles.
Contents of the Final Report
A GLP-compliant final report typically includes several key sections:
- Summary of methods and results: This section provides a concise description of the experimental design, materials used, and key findings. It allows reviewers to quickly understand how the study was conducted and what outcomes were observed.
- Statistical analyses: Where applicable, the report must include detailed statistical evaluation of the data. This may involve descriptive statistics, inferential tests, confidence intervals, and any models used to interpret results. The statistical methods used must be clearly described and justified.
- Discussion and conclusions: This section interprets the results in relation to the study objectives. It explains the significance of the findings, their implications, and any limitations of the study.
- Deviations from the protocol: Any changes or deviations from the original approved protocol must be clearly documented. This includes explanations of why the deviation occurred and an assessment of its potential impact on the study outcomes.
The report must be clear, logically structured, complete, and free from bias, ensuring that it can be independently reviewed and understood.
Archiving and Record Retention
GLP places strong emphasis on the secure archiving and long-term retention of all study-related materials. This requirement ensures that scientific data remain accessible for future verification, regulatory inspections, or re-analysis if necessary. All archived materials must be stored in a controlled, secure environment that prevents loss, deterioration, or unauthorized access. Proper indexing and cataloging systems are essential so that materials can be retrieved efficiently when required.
Types of Archived Materials
The scope of archiving under GLP is extensive and includes:
- Study protocols and final reports: These provide the official documentation of study design and outcomes.
- Raw data: Original observations, laboratory notebooks, electronic data files, and instrument outputs must be preserved in their unaltered form.
- Samples and specimens: Physical materials such as tissue samples, chemical residues, or biological specimens may be retained for reanalysis or verification.
- Standard Operating Procedures (SOPs) and training records: These documents demonstrate that appropriate procedures were followed and personnel were adequately trained.
Proper archiving ensures long-term data integrity and supports regulatory compliance. It allows studies to be reviewed or audited even years after completion, reinforcing transparency and accountability in scientific research.
Compliance and Regulatory Framework
GLP is not a voluntary system; it is a formally enforced regulatory framework adopted by many countries to ensure that non-clinical laboratory studies are conducted with a high level of integrity, traceability, and reliability. Regulatory compliance with GLP is often a mandatory requirement for studies that are submitted to government agencies for product registration, safety evaluation, or market approval. This is especially critical in sectors such as pharmaceuticals, pesticides, industrial chemicals, cosmetics, and veterinary products, where decisions directly impact human health and environmental safety. Several regulatory bodies oversee and enforce GLP standards at national and international levels. At the national level, agencies such as environmental protection authorities, food and drug administrations, and occupational safety institutions are responsible for inspecting laboratories and verifying compliance. These agencies ensure that laboratories adhere to standardized procedures and maintain proper documentation of all experimental activities.
At the international level, harmonization efforts are led by organizations that develop globally accepted GLP principles. These frameworks aim to ensure that data generated in one country is recognized and accepted in another, reducing the need for duplicate testing. This international alignment is particularly important for multinational companies seeking regulatory approval across multiple regions. Non-compliance with GLP principles can have serious consequences. Regulatory authorities may reject entire datasets if they are deemed unreliable or improperly documented, leading to delays in product approval or research progression. In more severe cases, laboratories may face legal action, financial penalties, or suspension of their operating licenses. Additionally, non-compliance significantly damages institutional credibility, reducing trust in both the laboratory and its scientific outputs. This loss of reputation can have long-term implications for funding, collaboration, and regulatory engagement.
Benefits and Limitations of GLP
The implementation of Good Laboratory Practice provides numerous advantages that contribute to the overall robustness of scientific research and regulatory decision-making. One of the most significant benefits is the enhancement of data quality and reliability. By enforcing standardized procedures, rigorous documentation, and quality assurance mechanisms, GLP ensures that experimental results are accurate, reproducible, and scientifically defensible.
Another major advantage is the facilitation of international acceptance of data. Because GLP principles are harmonized across many countries, data generated under compliant conditions in one region can often be accepted by regulatory authorities in another. This reduces duplication of studies, saving both time and financial resources while minimizing the use of animals and other experimental materials. GLP also plays a crucial role in protecting public health and the environment. By ensuring that safety assessments of chemicals, pharmaceuticals, and other substances are based on reliable data, regulatory agencies can make informed decisions that reduce risks to humans, animals, and ecosystems. This preventive function is a key reason why GLP is embedded in regulatory systems worldwide.
GLP is not without limitations. One of the primary challenges is that it can be resource-intensive and costly to implement. Laboratories must invest in trained personnel, quality systems, documentation infrastructure, and equipment maintenance, all of which require sustained financial and administrative support. Additionally, GLP tends to focus heavily on procedural compliance rather than scientific innovation. While this ensures consistency and reliability, it may sometimes limit flexibility in exploratory or highly novel research settings. The extensive documentation and administrative requirements can also introduce a significant burden on researchers, potentially slowing down experimental workflows. Despite these limitations, the overall benefits of GLP particularly in ensuring data integrity, regulatory acceptance, and public safety far outweigh its challenges, making it an essential component of modern laboratory science.
REFERENCES
Dubey, R. C. and Maheshwari, D. K. (2004). Practical Microbiology. S. Chand and Company LTD, New Delhi, India.
Garcia L.S (2010). Clinical Microbiology Procedures Handbook. Third edition. American Society of Microbiology Press, USA.
Garcia L.S (2014). Clinical Laboratory Management. First edition. American Society of Microbiology Press, USA.
Madigan M.T., Martinko J.M., Dunlap P.V and Clark D.P (2009). Brock Biology of Microorganisms, 12th edition. Pearson Benjamin Cummings Inc, USA.
Mahon C. R, Lehman D.C and Manuselis G (2011). Textbook of Diagnostic Microbiology. Fourth edition. Saunders Publishers, USA.
Prescott L.M., Harley J.P and Klein D.A (2005). Microbiology. 6th ed. McGraw Hill Publishers, USA. Pp. 296-299.
Ryan K, Ray C.G, Ahmed N, Drew W.L and Plorde J (2010). Sherris Medical Microbiology. Fifth edition. McGraw-Hill Publishers, USA.
Salyers A.A and Whitt D.D (2001). Microbiology: diversity, disease, and the environment. Fitzgerald Science Press Inc. Maryland, USA.
Discover more from Microbiology Class
Subscribe to get the latest posts sent to your email.